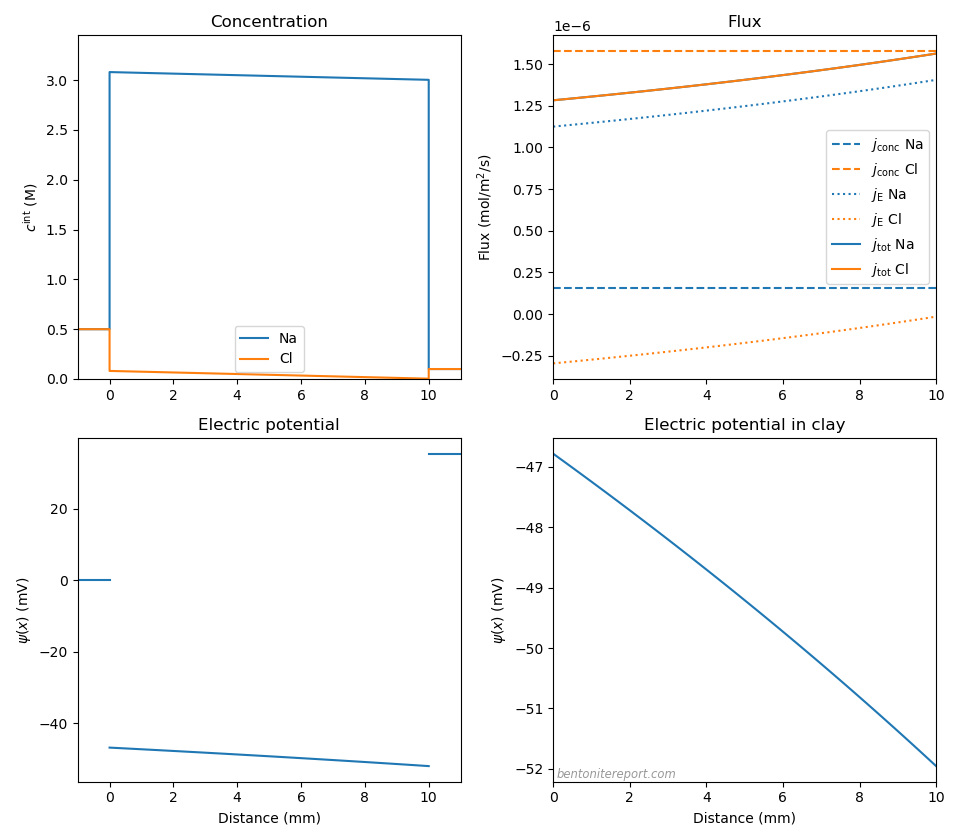
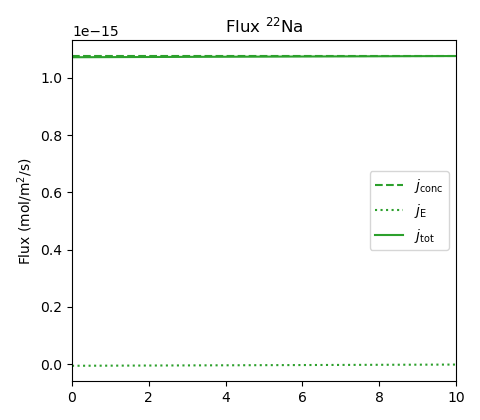
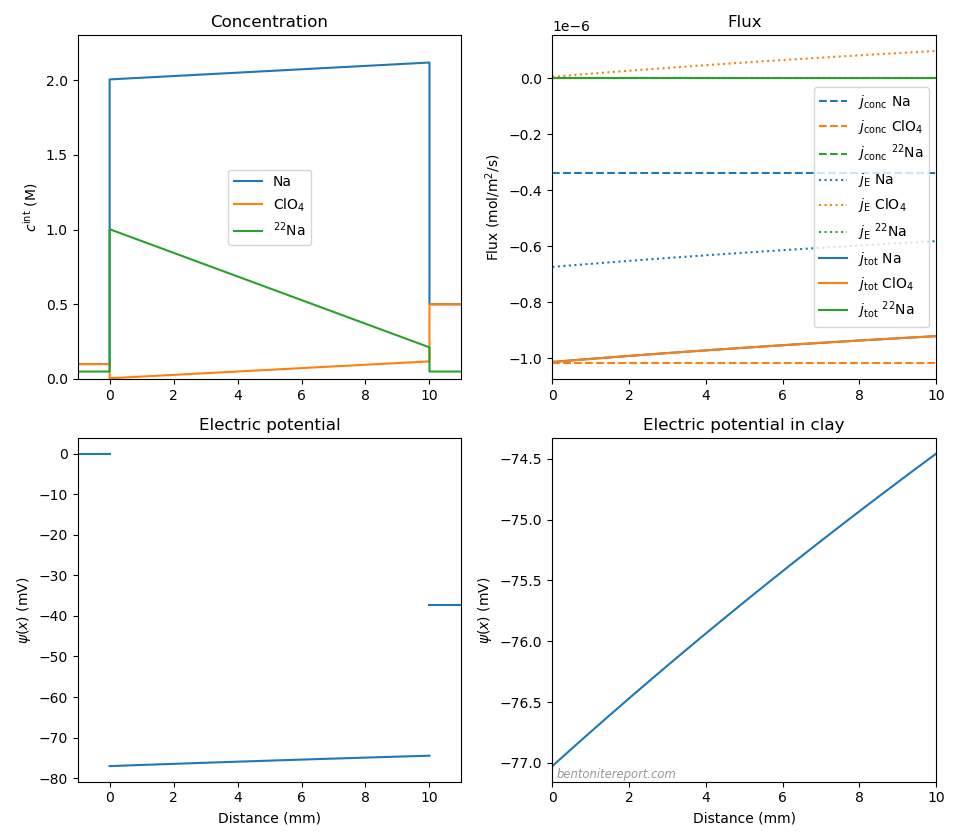
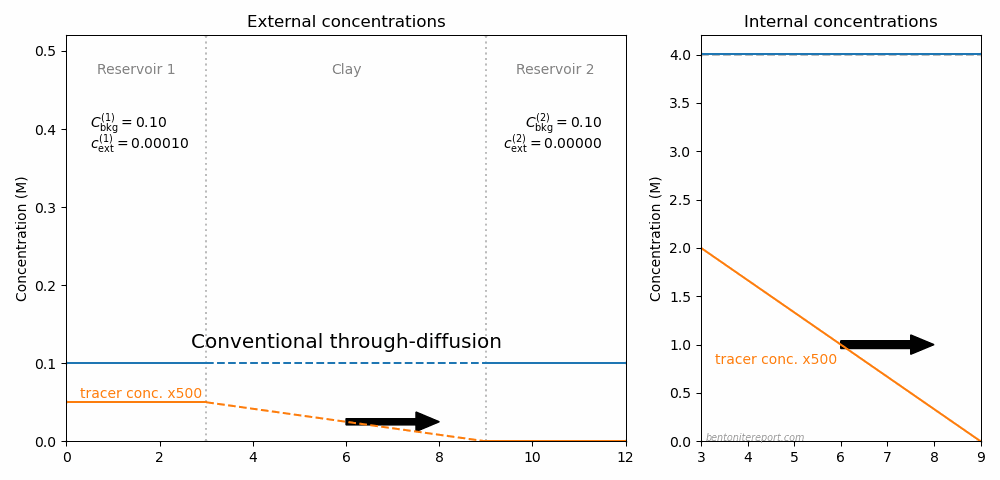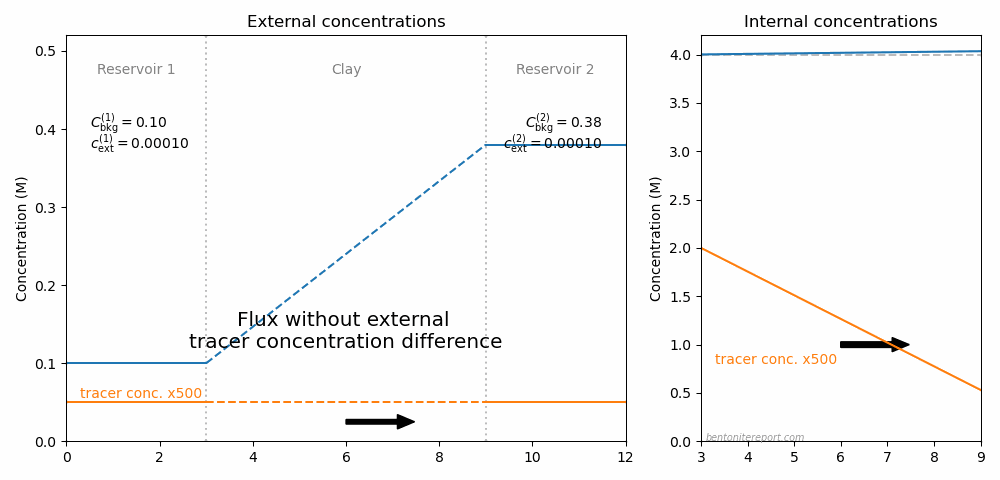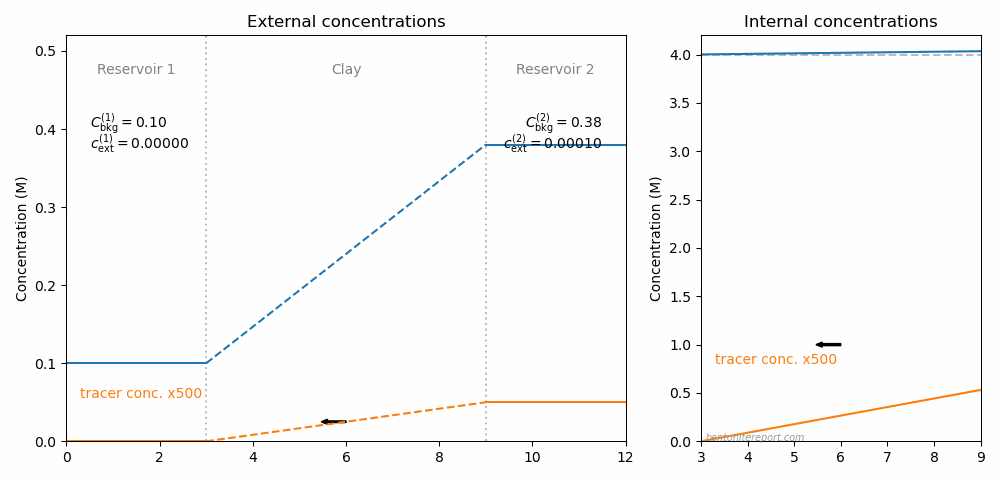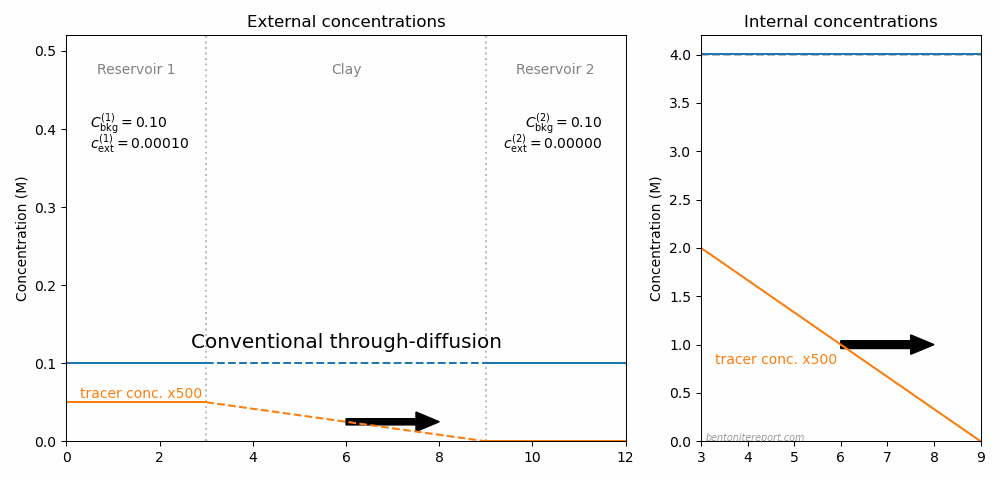
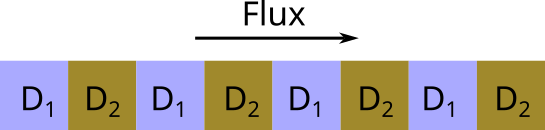We have discussed the seeming steady-state uphill testfrequently on the blog. What I find important to clarify is that the “uphill” effect is governed by the exact same mechanism that controls the behavior of cation diffusion in conventional through-diffusion tests: Donnan equilibrium.1 This insight seems to be not at all realized in the contemporary bentonite scientific literature.
With a tool available that solves the Nernst-Planck equations in bentonite, I would here like to present a complete2 modeling of the “uphill” test, with a direct comparison with the experimental results. It may also be compared with the previous evaluation, which only considered Donnan equilibrium and imposed linear concentration profiles, rather than solving the full transport equations.
Set-up
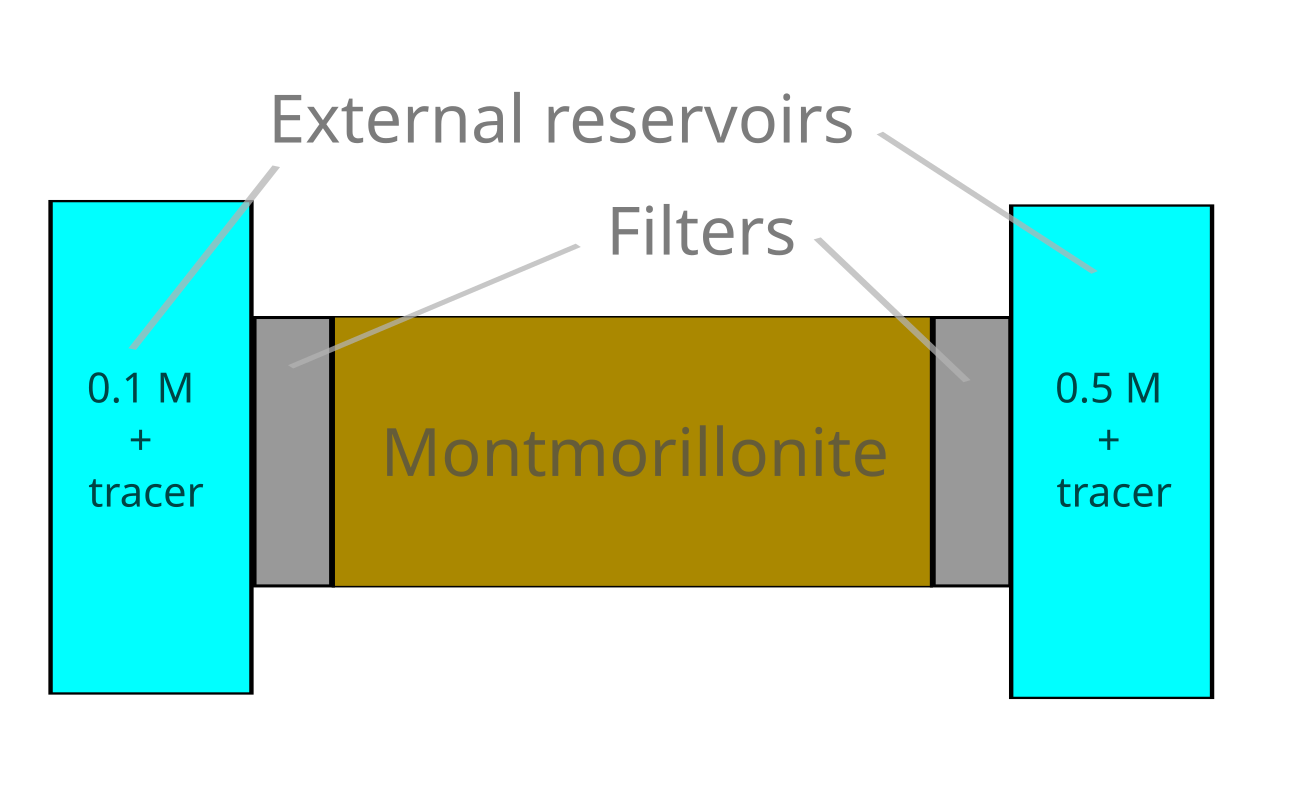
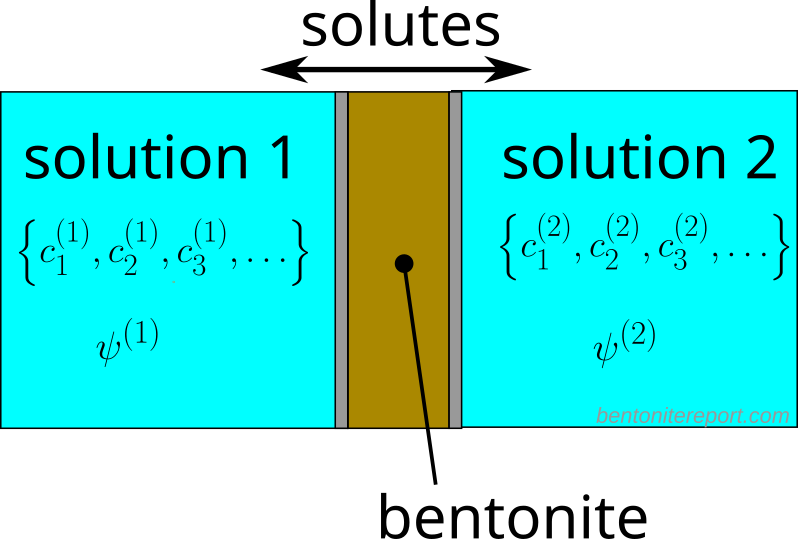
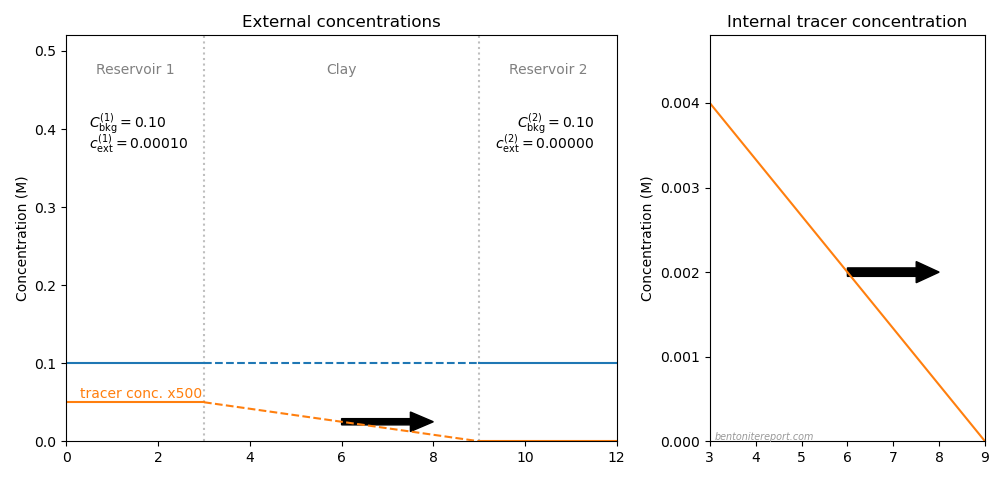
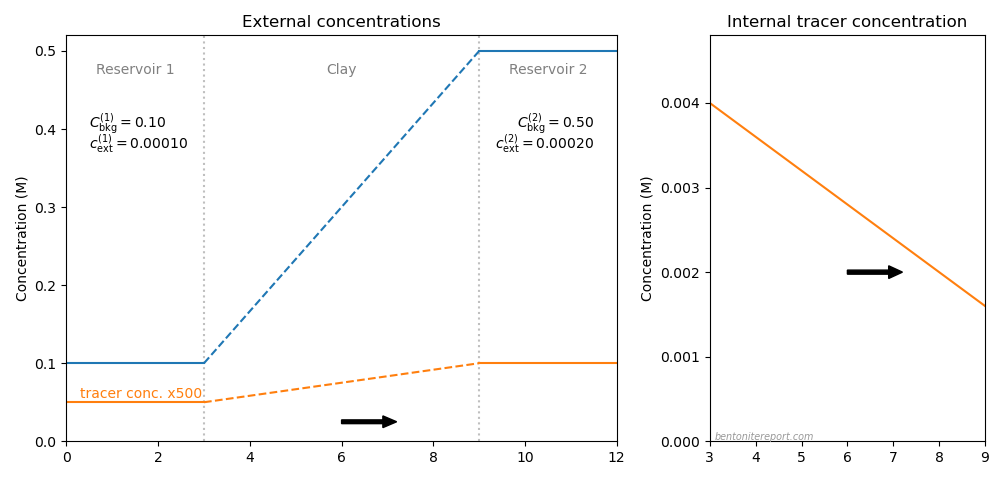
As is evident from the results of the modeling, and what we already have concluded on several occasions, the inclusion of multi-component diffusion actually makes no essential difference for the migration of the tracer in the “uphill” test. In contrast, it is crucial to include transport in the confining filters, which is to be excepted, since the test is performed with very high density montmorillonite contacted with relatively low external concentration. Likewise, the model must account for the varying concentration in the external reservoirs. Schematically the test set-up/model look like this
The (1D) Nernst-Planck equations are solved in the filters and the clay, and the clay is treated as a single domain with the homogeneous mixture model. Donnan equilibrium is maintained at the filter/clay interfaces.
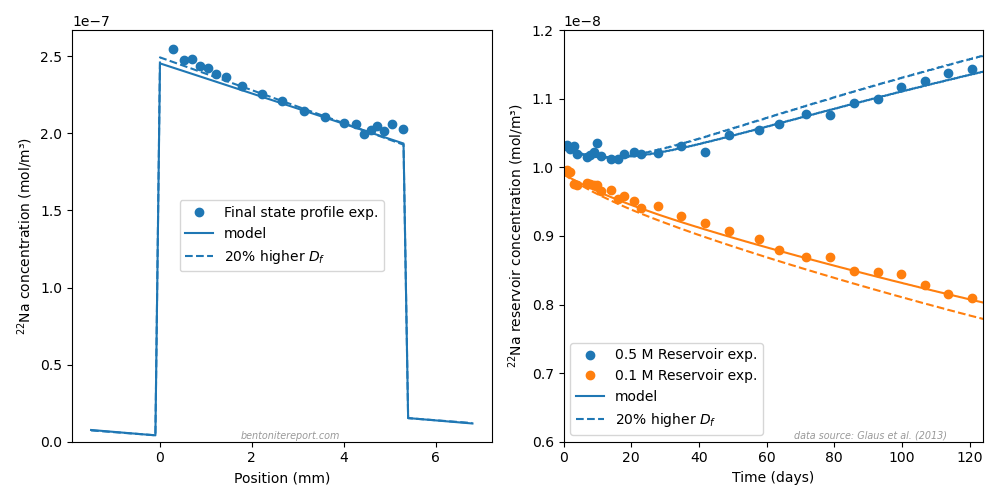
The “uphill” study (Glaus et al., 2013) presents two separate tests, where a sodium tracer diffuses in pure Na-montmorillonite samples of either 5.4 mm3 or 10 mm. Otherwise, the conditions are essentially identical. The left hand side reservoirs are kept at 0.1 M (NaClO4), and the right hand side reservoirs at 0.5 M (NaClO4). The nominal initial 22Na tracer concentration is 10-11 M in both reservoirs. The reservoir volumes are 100 ml for the test with a 10 mm sample and 250 ml for the 5.4 mm test.
Results
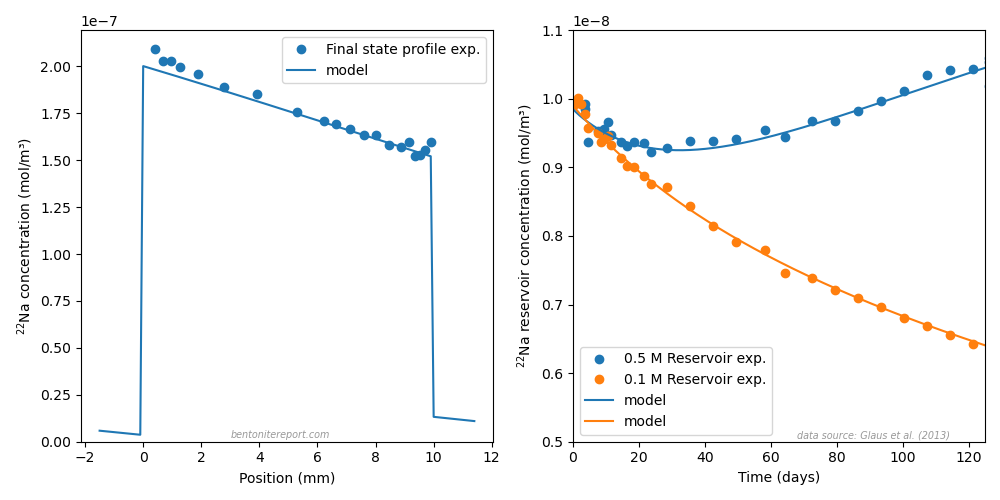
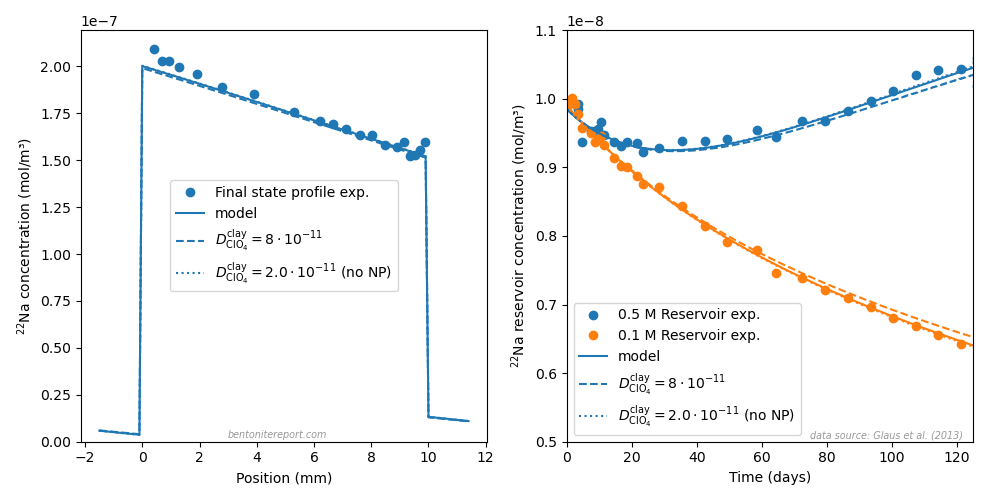
Here is a comparison between model and experiment for the 10 mm sample, for the final state concentration profile (after 162 days) and the evolution of the tracer concentration in the two reservoirs
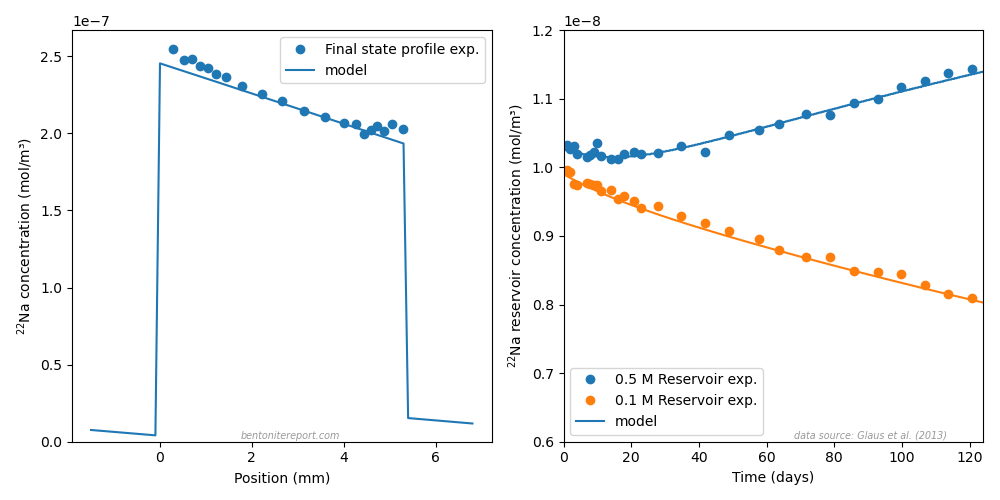
The corresponding plot for the 5.4 mm sample looks like this
The model parameters are
Sample
10 mm
5.4 mm
Porosity
0.321
0.321
\(c_\mathrm{IL}\) (CEC)
5.64 M (0.954 eq/kg)
6.21 M (1.05 eq/kg)
Cross section area
5.15 cm2
5.15 cm2
\(D_\mathrm{Na}^\mathrm{clay}\)
2.0⋅10-11 m2/s
1.9⋅10-11 m2/s
\(D_\mathrm{ClO_4}^\mathrm{clay}\)
2.67⋅10-11 m2/s
2.53⋅10-11 m2/s
Filter length
1.5 mm
1.5 mm
Filter porosity
0.321
0.321
\(D_\mathrm{Na}^\mathrm{filter}\)
6.0⋅10-11 m2/s
7.4⋅10-11 m2/s
\(D_\mathrm{ClO_4}^\mathrm{filter}\)
8.0⋅10-11 m2/s
9.9⋅10-11 m2/s
Volume reservoirs
100 ml
250 ml
\(c^\mathrm{res1}_{tr}\) (initial)
0.99⋅10-8 mol/m3
0.99⋅10-8 mol/m3
\(c^\mathrm{res2}_{tr}\) (initial)
0.985⋅10-8 mol/m3
1.03⋅10-8 mol/m3
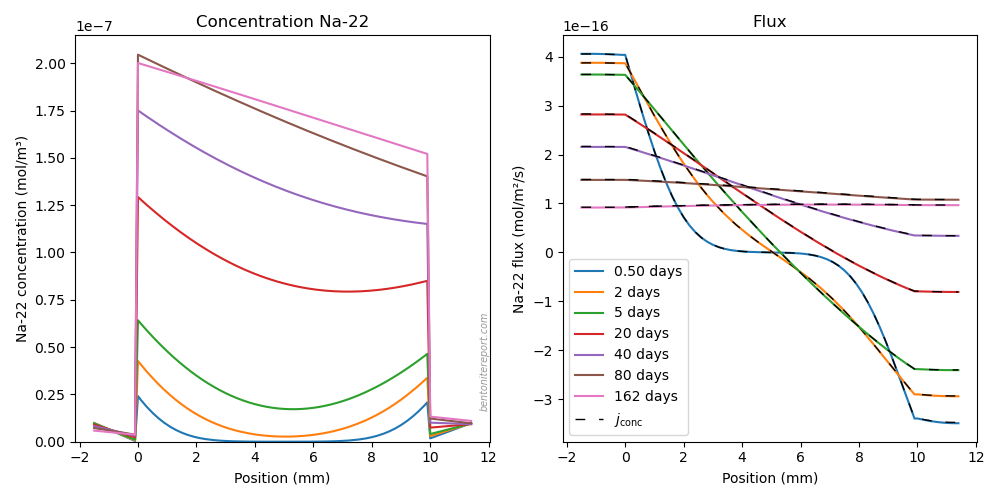
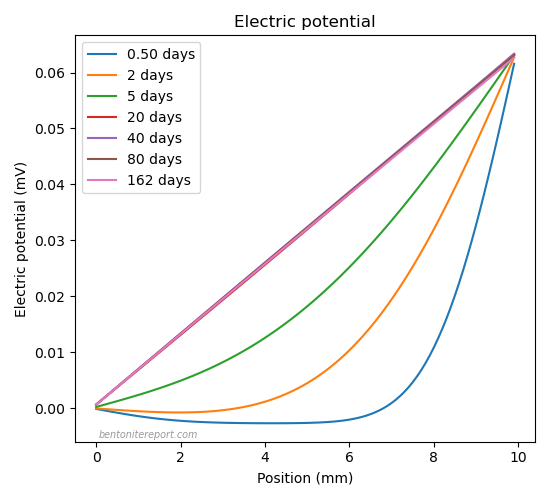
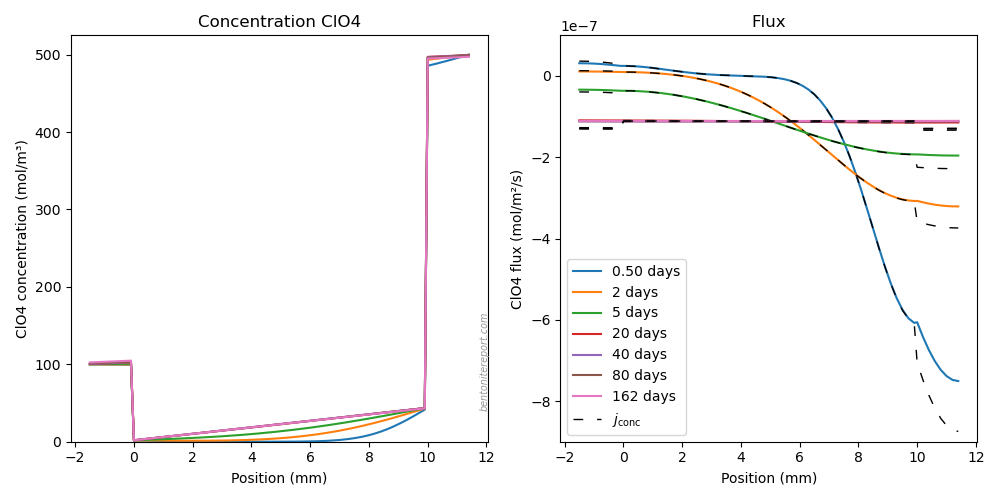
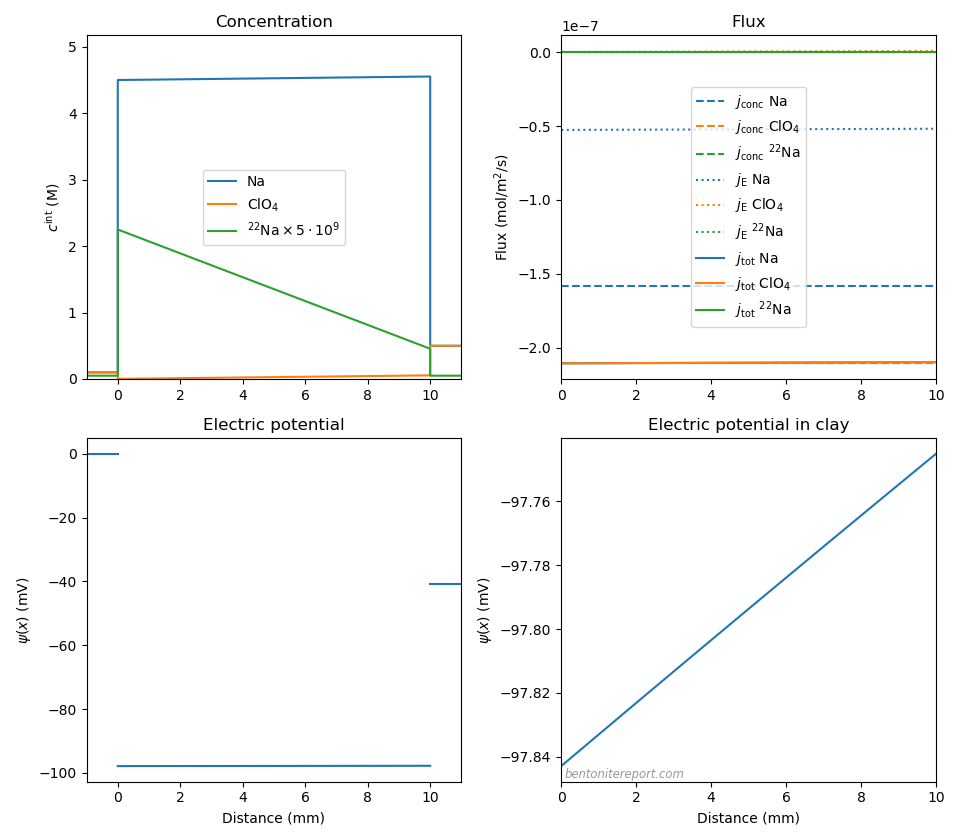
With the full model at our disposal, we can further investigate the evolution of the systems during the course of the tests. Below are plotted snapshots of tracer concentration, electric potential, and flux profiles in the 10 mm sample.
The flux snapshots show both the full flux (concentration gradient + electromigration terms) and the “Fickian” flux, i.e. the part of the flux driven by the tracer concentration gradient (dashed black lines). As in the previous assessment, we note that electromigration effects on the tracer flux is completely negligible; the flux is essentially completely “Fickian”. The diffusion potential — defined as the steady-state electric potential difference due to electromigration across the clay — is 0.06 mV.4 This number is even smaller than in the previous evaluation (0.09 mV), because the parameter \(c_\mathrm{IL}\) is larger here (here, \(c_\mathrm{IL}\) was used as a fitting parameter, as further discussed below).
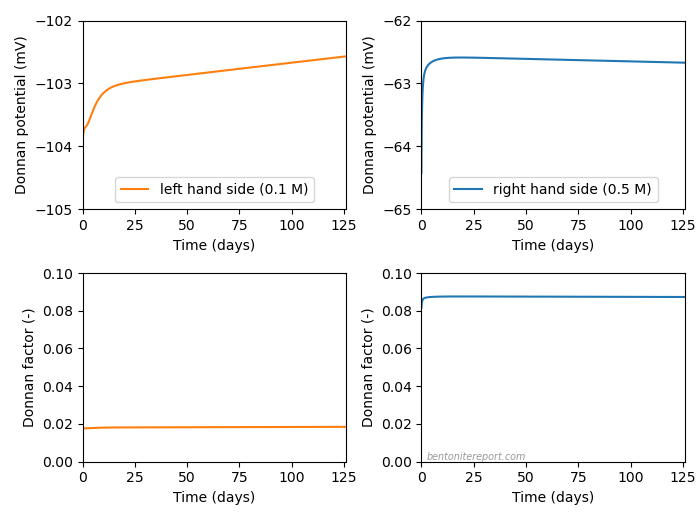
The evolution of the Donnan potentials, and corresponding Donnan factors, at the two clay/filter interfaces look like this
As the reservoirs have finite size, their compositions slowly change during the course of the test, and a corresponding change is seen in the Donnan potentials. As the concentration is lower in the left hand side reservoir, the change is more pronounced there. The effect on the Donnan factors (whose inverses determine the enhancement of the tracer concentrations in the clay) is nevertheless small. The contribution from Donnan equilibrium to the membrane potential — which corresponds to the difference between these Donnan potentials — evolves in the range 40.9 — 39.9 mV, in reasonable agreement with the previously evaluated (constant) contribution of 41.1 mV.5
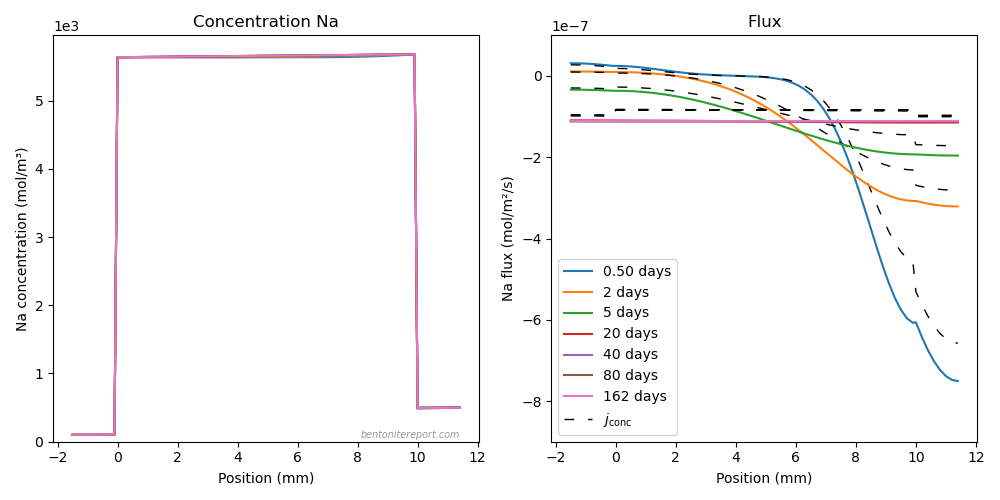
The evolution of the main salt behaves very differently as compared with the tracer
Most notably, the flux of stable NaClO4 is directed opposite to that of the tracers (“downhill”). Note also that this flux is not limited by filter diffusivity (the concentration drop across the filters is negligible) and that this process reaches (quasi-)steady-state within 20 days. The reduced internal concentration gradient, as compared with the external concentration difference, explains why the process of equilibrating the main salt between the reservoirs is much slower in comparison to the “uphill” process. This, in turn, is of course also a consequence of Donnan equilibrium.
An electromigration effect is observed for both Na and ClO4 in the filters (ClO4 is beeing slowed down, while Na is boosted). In the clay (\(0 \le x \le 10\)), electromigration is mostly noticeable for sodium, because this ion has a much larger concentration there.
Fitting strategy and sensitivity analysis
We note that essentially perfect agreement can be achieved between model and experiments. But we note also that the model results depend on a quite large set of parameters. Some of these are quite easily constrained by the experiment itself, while we have used others as fitting parameters.
Regarding sample geometry, we have simply chosen the nominal density (1.9 g/cm3) and corresponding porosity (assuming a grain density of \(\rho_s\) = 2.8 g/cm3). Likewise, sample lengths and cross sectional area and reservoir volumes are fixed by what is stated in the paper.3 The filter porosity is chosen to be equal to the clay porosity, and we have chosen a filter length of 1.5 mm (as stated in Glaus et al. (2007)).
We have furthermore chosen to keep the same ratio (0.75) as for bulk solutions between sodium and perchlorate diffusivities in the montmorillonite.
The main factors that have been varied are the parameter \(c_\mathrm{IL}\) (which can be considered equivalent with varying the cation exchange capacity), the sodium diffusivity in clay (\(D_\mathrm{Na}^\mathrm{clay}\)) and filters (\(D_\mathrm{Na}^\mathrm{filter}\)), and the initial reservoir tracer concentrations (\(c_\mathrm{tr}^\mathrm{res1}\), \(c_\mathrm{tr}^\mathrm{res2}\)). For the initial tracer concentrations, it should be straightforward to accept the chosen values as evidenced by the outflux-plots. We will not comment them further.
For the sodium diffusivity in the clay, the experimental results already give a quite good hint of its value, since we can evaluate both a (quasi-)steady-state flux and a corresponding internal tracer concentration gradient. Such evaluations are naturally burdened by uncertainties, but gives a rough estimate for the sodium diffusivity of \(\sim\)2⋅10-11 m2/s for both tests. Such a value is also expected from previous tests of sodium diffusivity in sodium montmorillonite. In particular, Glaus et al. (2007) evaluate diffusivities in nearly identical samples in the range 1.6⋅10-11 m2/s — 2.2⋅10-11 m2/s. Also Kozaki et al. (2005) report diffusivities of this size, albeit in a different type of sodium montmorillonite material (“Kunipia-F”).
With the clay diffusivity quite constrained there is not much choice left but to fit the model by varying the filter diffusivity (and \(c_\mathrm{IL}\)). This is the fitting strategy chosen here. In the plot below is shown the high sensitivity to the filter diffusivity in an alternative model where the filter diffusivity is increased by 20%, in the 5.4 mm sample.
Note that the transport capacity of the filter is set by a combination of its porosity, length and diffusivity (\(\phi D/L\)). As we have no exact information on porosity and length (but keep them fixed), the filter diffusivity variation should be viewed as a variation of the filter transport capacity as a whole.
We also explore the (in)sensitivity of the multi-component aspect of the model, by investigating alternative models where the Na/ClO4 diffusivity ratio is varied in the clay. The following plot demonstrates how insignificant the Nernst-Planck aspect is for the tracer, by comparing a model where the perchlorate diffusivity is increased to be a factor 4 larger than the sodium concentration, and a model where all diffusivities are put equal (thus eliminating electromigration completely).
The present modeling could consequently have been done using only Fick’s laws, while giving essentially the same result (for the tracer).
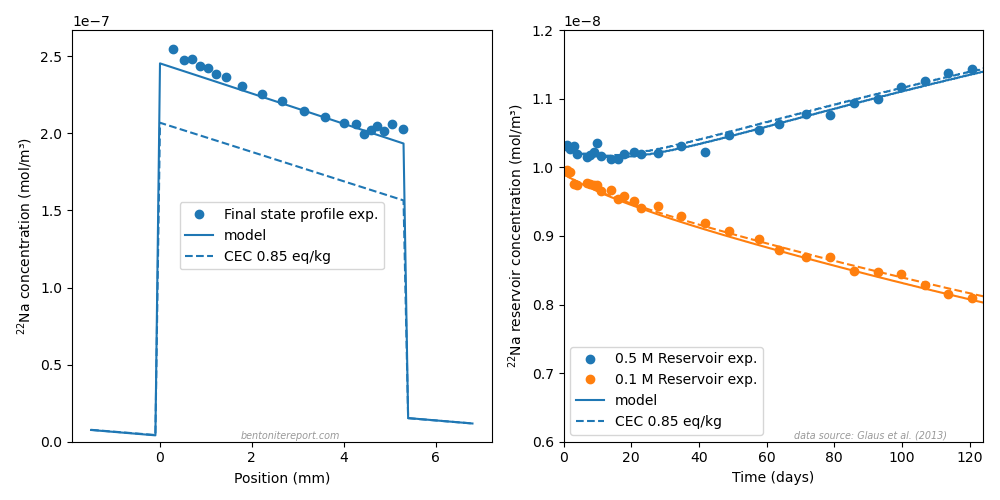
The only “anomalous” parameter choice that was made in the model fittings is an unjustifiably large value of the cation exchange capacity (or \(c_\mathrm{IL}\)). The reported cation exchange capacity for the material (“montmorillonite from Milos”) is 0.8 — 0.88 eq/kg, while the fitted \(c_\mathrm{IL}\) values correspond to values approximately 10 — 30% larger. Variations of this parameter directly affects the absolute value of the internal concentration profile, but is quite insensitive to the resulting outfluxes, as seen here in a variation of the 5.4 mm model using a tabulated value for the CEC (0.85 meq/kg, \(c_\mathrm{IL}\) = 5.02 M).
This mismatch may indicate that some effect or process is missing or not properly handled in the models. However, we should keep in mind that we have assumed the nominal density when converting the reported tracer concentration profiles (which have unit mol/kg dry clay) to pore concentrations (mol/m3). At these very high densities, \(c_\mathrm{IL}\) is hyper sensitive to the exact water-to-solid mass ratio, \(w\) (\(c_\mathrm{IL}\) depends on the inverse of \(w\)). An actual density of the samples somewhat lower than the nominal value may thereby explain the discrepancy. We may also note that the fitted \(c_\mathrm{IL}\) value varies significantly for the two tests, which also may indicate that this mismatch is a matter of experimental uncertainty, rather than being an indication of an actual effect. A further investigation of this possible anomaly would be much helped by results from additional tests.
Conclusions
The fundamental mechanism behind the seeming uphill diffusion effect is Donnan equilibrium, just as for conventional cation through-diffusion.
Effects of electromigration for the tracer is negligible, even though non-negligible concentration differences are maintained across the clay (and, consequently, multi-component diffusion is active).
Donnan equilibrium also explains the negligible transport of the main electrolyte; the time scale for equilibrating the main salt across the reservoirs is much larger then the time scale of the tests.
Filter transport limitations strongly influence the overall process.
The only possible unexplained feature of the modeling is an indication of a too large cation exchange capacity required to perfectly fit the data (an approximate 10 — 30% increase).
As I have whined about before, I think it is a pity that only a single study exists that explores this effect. More empirical input — e.g. for lower densities, other concentration configurations, other ion types, and membrane potentials — will greatly help process understanding.
Footnotes
[1] Someone might perhaps object to this and claim that the relevant mechanism is ion exchange. But ion exchange is nothing but a manifestation of a system that is progressing towards Donnan equilibrium. And any talk in this context about “sorption”, “sites”, surface complexation, Stern layers, immobilization, etc., is just that: talk.
[2] It’s of course dangerous to call a model “complete”. In it current state, the model still ignores e.g. pressure (but is at least in principle able to describe swelling).
[3] The nominal size of one sample is 5 mm, but it is clear from the concentration profile presented in Glaus et al. (2013) that it is slightly longer than this. In the model, we have adopted the length 5.4 mm and will here refer to it as the 5.4 mm sample.
[4] Interestingly (perhaps), the potential drop across the filters are actually comparable with that across the clay.
[5] As the Donnan potentials does not stay completely constant, we have set the reference such that \(\psi(0) = 0\) in the the plot showing the evolution of the electric potential in the clay. It would otherwise be hard to visualize, since its variation is so small.
I have started developing a tool that numerically solves the Nernst-Planck equations in bentonite on a 1D grid. It treats the bentonite within the homogeneous mixture model, accounting for the structural clay charge via the parameter \(c_\mathrm{IL}\). It also handles external solution reservoirs of finite size, and Donnan equilibrium is maintained dynamically at the interfaces between clay and external solutions. At the moment, the tool does not account for activity coefficients.
Here \(D_c\), \(z\) and \(c^\mathrm{int}\) are the diffusion coefficient, charge number and concentration, respectively, for the considered species. \(\phi\) denotes porosity. For convenience, we denote the spatial derivative (\(\partial/\partial x\)) by \(\nabla\). \(\tilde{\psi}\) is the dimensionless electric potential,1 whose gradient is
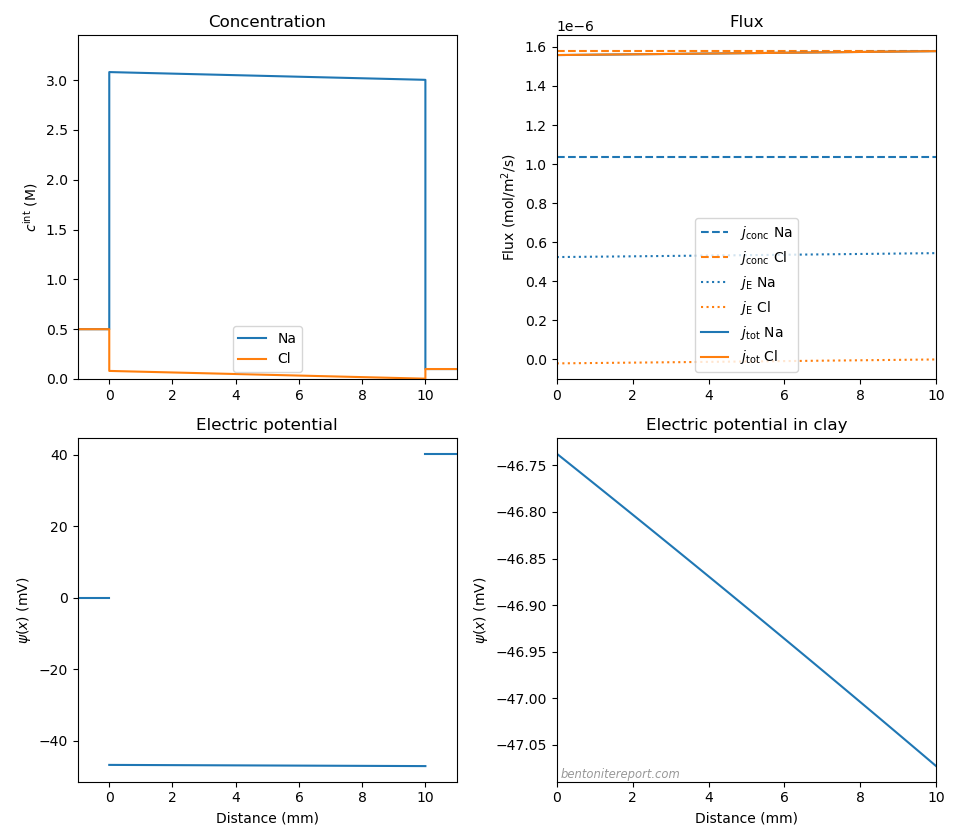
As we noted previously, compacted bentonite often only show minor effects of electromigration, with diffusion potentials below 1 mV. This is because substantial counter-ion concentrations reduce the size of the electric field required to counteract effects from differences in ion mobilities (the denominator in eq. 4 is always large). To see some effect when exploring the full solution of the Nernst-Planck equation, let’s therefore focus on one of the hypothetical, “extreme” cases considered in the previous post. This test case considers diffusion of CaCl2 through bentonite of quite low density (\(c_\mathrm{IL}\) = 1.5 M), with a 10 times lower calcium mobility relative to chloride, and has a 0.25 M CaCl2 reservoir on the left side and a 0.05 M CaCl2 reservoir on the right side.
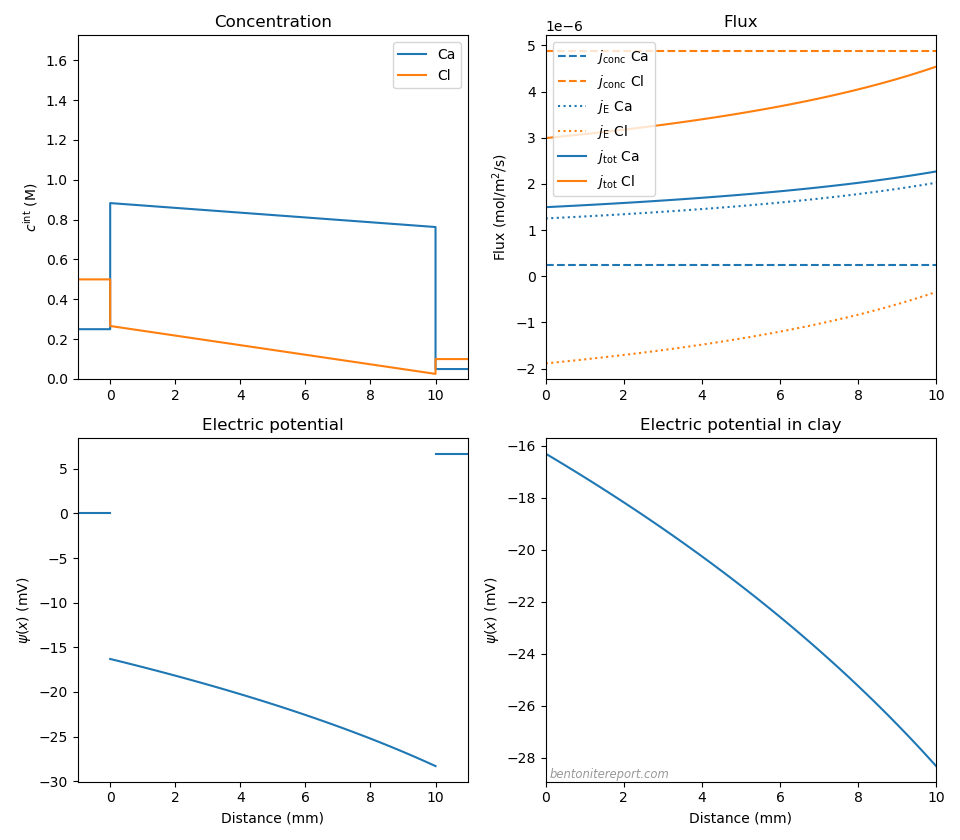
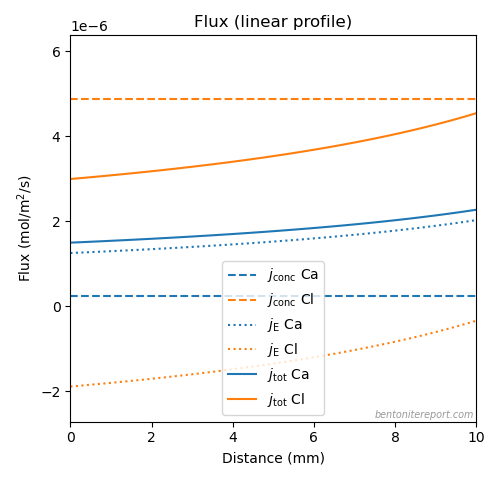
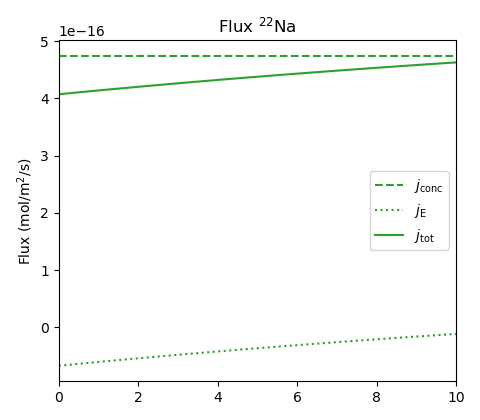
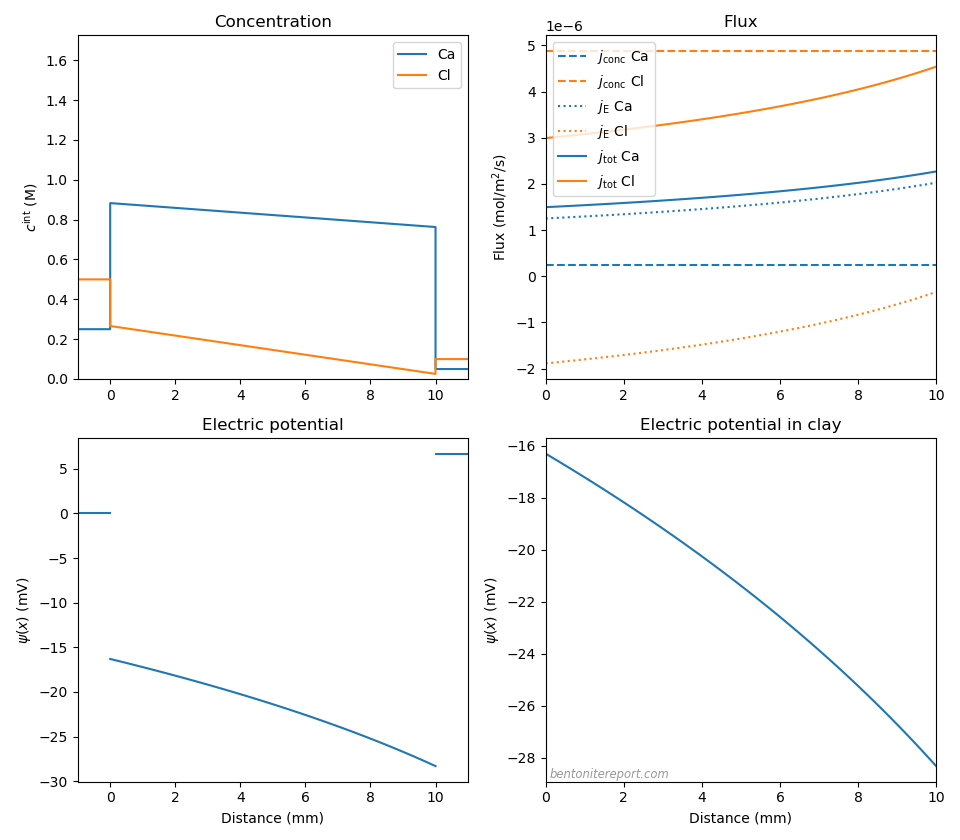
In the previous consideration we did not solve the full Nernst-Planck problem (eqs. 1— 5), but imposed a linear concentration gradient across the clay and calculated the corresponding electric potential (eq. 4) and contributions to the flux (eqs. 2 and 3). For the “extreme” CaCl2-case, the results of this approach looked like this (relevant model parameters are summarized below the figures)
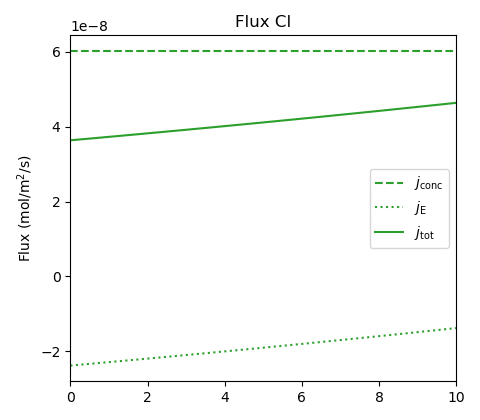
Model parameters: \(c_\mathrm{IL}\)= 1.5 M, \(D_\mathrm{Ca}^\mathrm{clay}\) = 0.203⋅10-10 m2/s, \(D_\mathrm{Cl}^\mathrm{clay}\) = 2.03⋅10-10 m2/s, c(1) = 0.25 M CaCl2,c(2) = 0.05 M CaCl2.
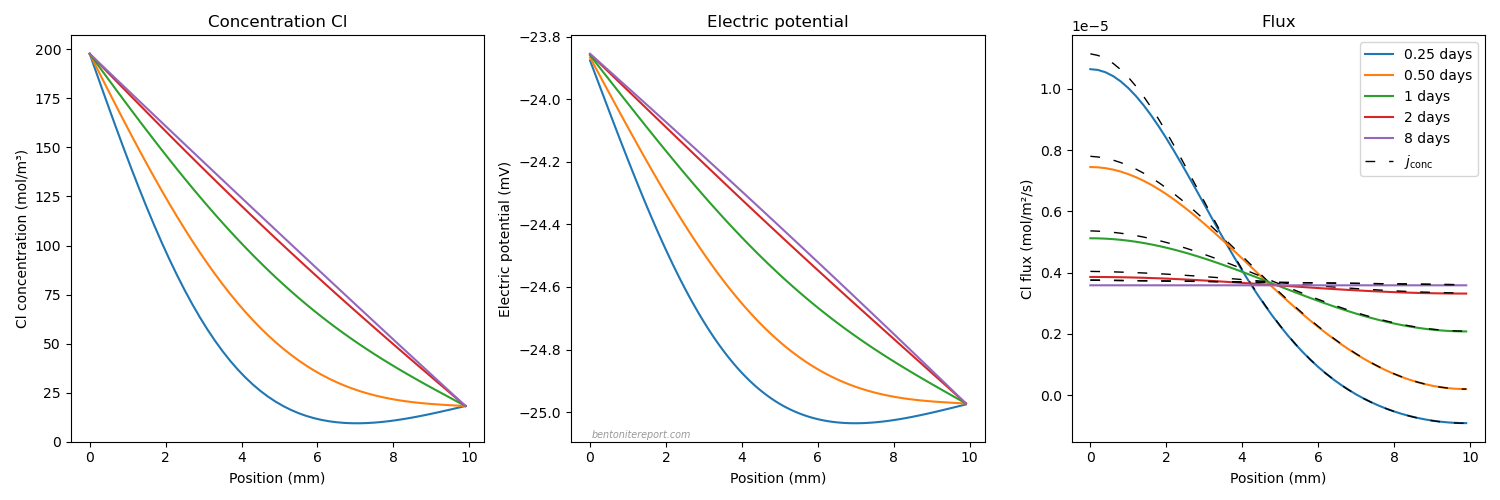
The concentration gradient contributions to the fluxes are constant throughout the clay domain (dashed lines), signifying the imposed linear concentration profile. In contrast, the total fluxes varies significantly (full lines), indicating that the system is not in steady-state. As the calcium mobility is 10 times lower than the chloride mobility, it is no surprise that the calcium flux is dominated by the electromigration contribution (dotted lines). Also the chloride flux is seen to be significantly influenced (retarded) by the electric field. The diffusion potential was evaluated to -12.1 mV, while the Donnan contribution to the membrane potential is +18.8 mV, giving an evaluated membrane potential of +6.7 mV (for a fuller discussion on these potentials, see the previous post). But note that this “membrane potential” is not completely accurately evaluated, as the linear concentration profile does not correspond to steady-state.
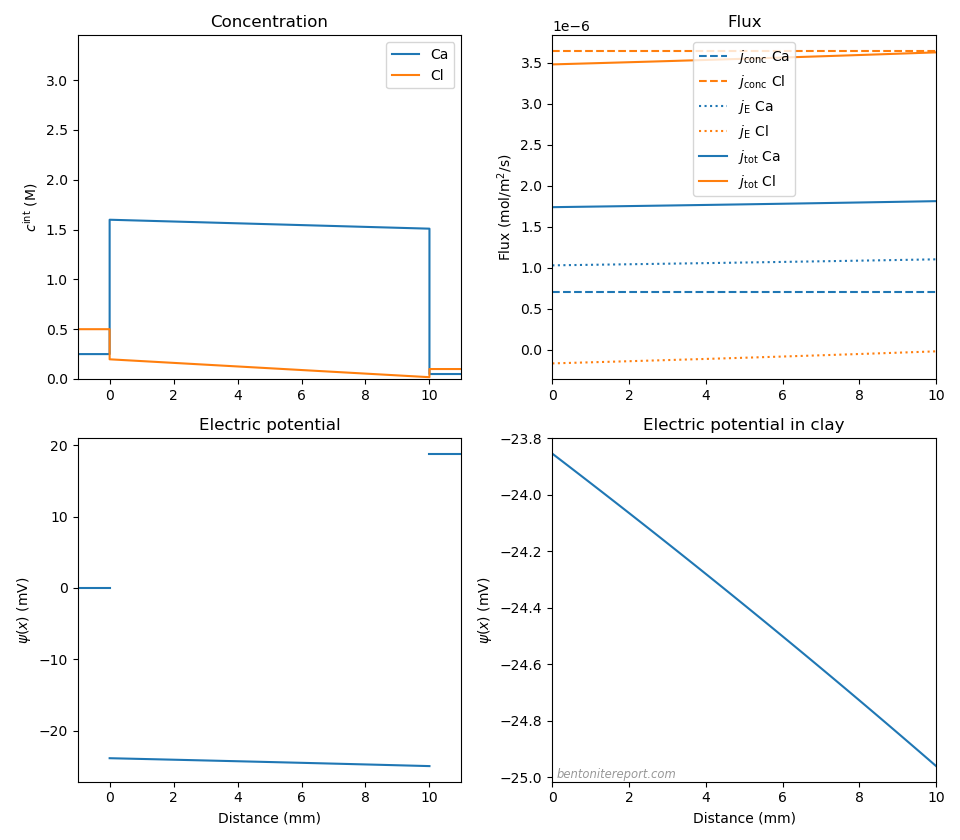
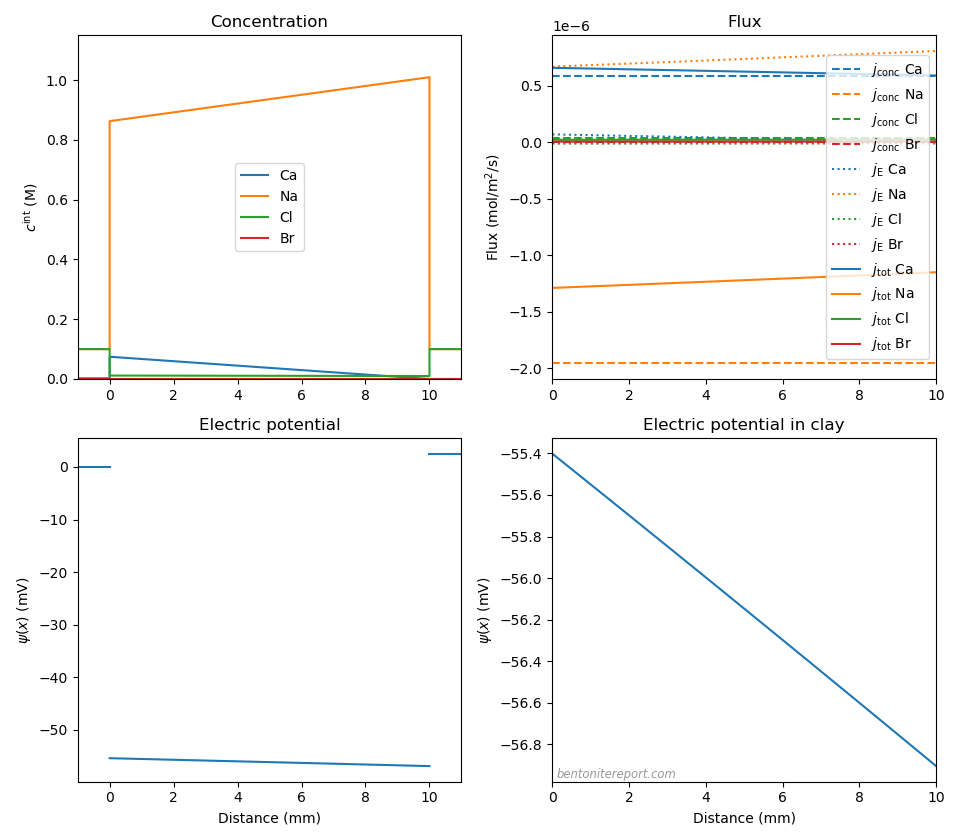
By solving the full Nernst-Planck framework (eqs. 1— 5), we can investigate the size of the error in the above treatment. The full solution looks like this
Model parameters: \(c_\mathrm{IL}\) = 1.5 M, \(D_\mathrm{Ca}^\mathrm{clay}\) = 0.203⋅10-10 m2/s, \(D_\mathrm{Cl}^\mathrm{clay}\) = 2.03⋅10-10 m2/s, c(1) = 0.25 M CaCl2,c(2) = 0.05 M CaCl2.
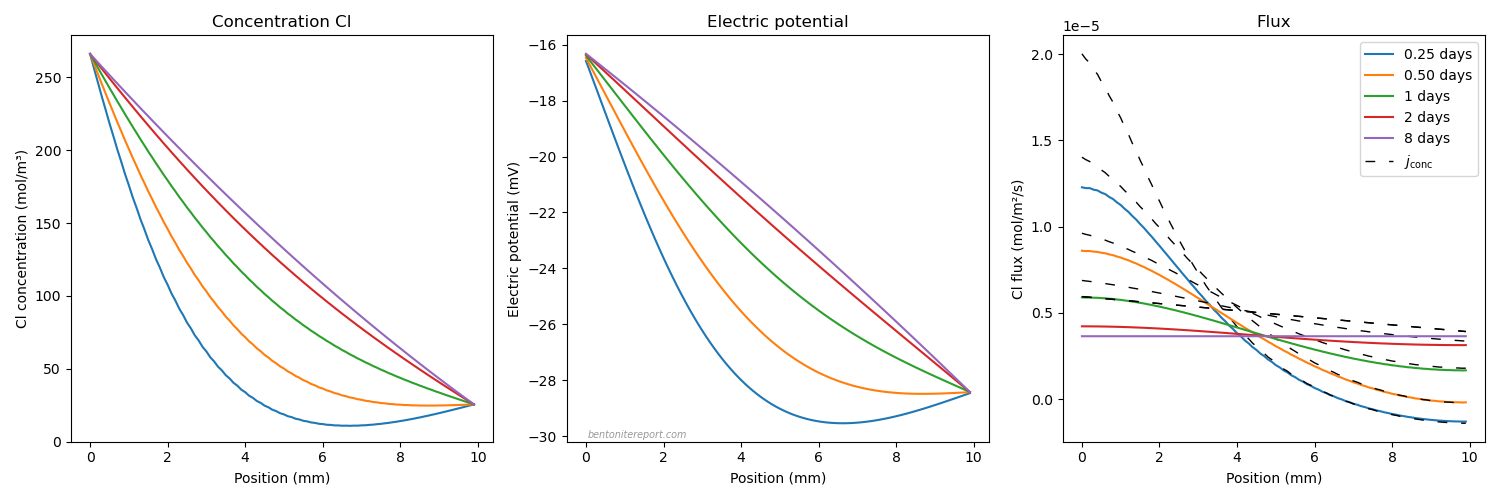
This figure shows snapshots of profiles of the chloride concentration, the electric potential and the chloride flux at 0.25, 0.5, 1, 2 and 8 days; the initial condition is a chloride free calcium clay. In the flux diagram are plotted both the full flux (solid lines) and the concentration gradient contribution3 (dashed lines). Although only barely noticeable, the steady-state has a slightly non-linear concentration profile, resulting in a varying “Fickian” flux contribution (\(j_\mathrm{conc}\)). However, compared with the case with imposed linear concentration profiles, the diffusion potential changes only slightly, to -12.2 mV, giving a true steady-state membrane potential of +6.6 mV.
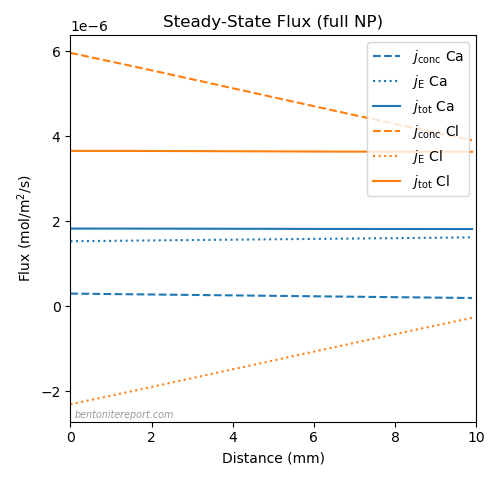
Below are compared the steady-state fluxes for both calcium and chloride with the fluxes evaluated from imposing linear concentration profiles.
The — now constant — total Cl flux is 3.64⋅10-6 mol/m2/s, to be compared with a variation between 3.0⋅10-6 mol/m2/s and 4.5⋅10-6 mol/m2/s in the previous calculation. Likewise, the previously constant chloride concentration gradient flux of 4.88⋅10-6 mol/m2/s now varies between approximately 6⋅10-6 mol/m2/s and 4⋅10-6 mol/m2/s.
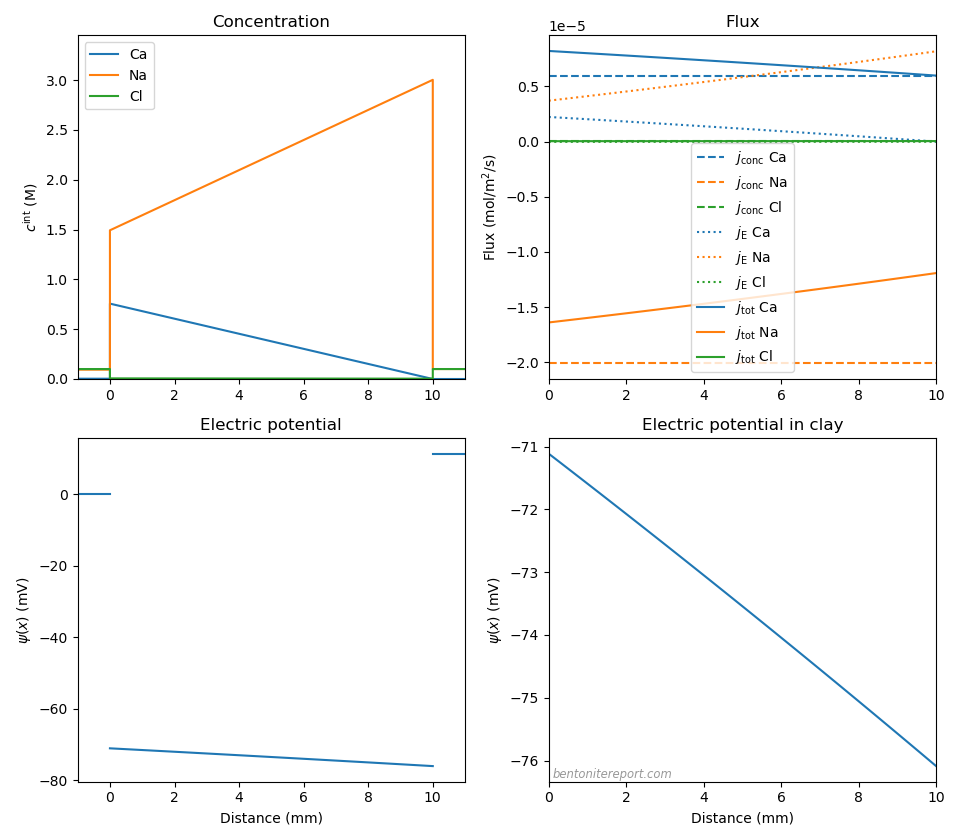
We should remind ourselves that we have considered an “extreme” case, and also take a look at a more realistic/relevant model, to illustrate the often quite insignificant contribution of electromigration in bentonite. In the plot below we are solving the full Nernst-Planck framework for CaCl2 through diffusion, as previously, but now with a denser clay (\(c_\mathrm{IL}\) = 3.0 M) and by setting the Cl/Ca diffusivity ratio equal to the corresponding value in bulk water (2.57).
Model parameters: \(c_\mathrm{IL}\) = 3.0 M, \(D_\mathrm{Ca}^\mathrm{clay}\) = 0.79⋅10-10 m2/s, \(D_\mathrm{Cl}^\mathrm{clay}\) = 2.03⋅10-10 m2/s, c(1) = 0.25 M CaCl2,c(2) = 0.05 M CaCl2.
We considered also this case in the previous treatment. The diffusion potential is here -1.1 mV, and the membrane potential +18.8 mV, which we also evaluated previously. Also the steady-state fluxes are essentially the same as previously evaluated: the chloride steady-state flux is 3.6⋅10-6 mol/m2/s, while in the case with an imposed linear concentration profile, the total flux varies between 3.5⋅10-6 mol/m2/s and 3.6⋅10-6 mol/m2/s. All of these similarities are of course a manifestation of that the linear concentration profiles are very close to being the true steady-state profiles.
Note how similar the concentration gradient contribution is to the total flux for chloride during the entire process, in contrast to the behavior in the “extreme” case. This indicates that chloride here essentially show “Fickian” diffusion, at the rate set by its individual diffusion coefficient. It should be remembered, however, that the full transport process is still influenced by electromigration, since charge neutrality is required (the slower calcium ions are “boosted” by the electric field). But such effects are in principle no different from how salts generally diffuse in ordinary solutions.
In the original publication, these examples consider a clay component consisting of parallel bulk water and “diffuse layer” domains, and linear concentration profiles are imposed in the bulk water; a main NaCl concentration decreases from 0.1 M to 0.001 M in both examples, while different 22Na and 36Cl concentrations are maintained. In “example 2”, the tracer concentrations decrease in proportion with the main bulk electrolyte, from 10-9 M to 10-11 M. In “example 3” the bulk water tracer concentrations are kept at a constant value 10-9 M.
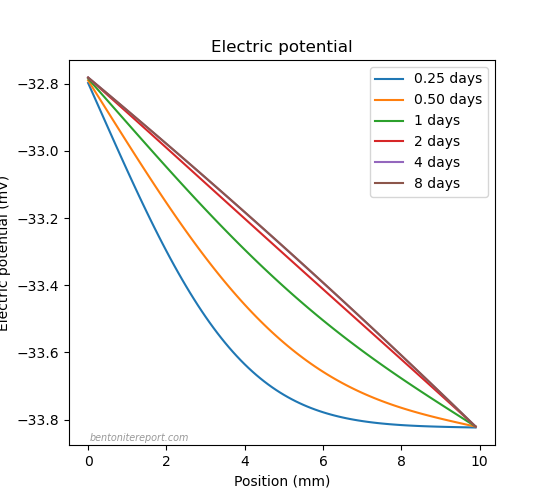
Here we instead consider only the “diffuse layer” domain, and impose the bulk water concentrations as boundary conditions. The “diffuse layer” domain is characterized by the parameter \(c_\mathrm{IL}\) = 0.33 M (Tournassat and Steefel (2015) call this quantity \(q\)) and is assumed initially free of chloride and tracers. Solving the full Nernst-Planck equations gives the following evolution of the electric potential (this is the same for both examples)
Model parameters: \(c_\mathrm{IL}\)= 0.33 M, \(D_\mathrm{Na}^\mathrm{clay}\) = 1.33⋅10-10 m2/s, \(D_\mathrm{Cl}^\mathrm{clay}\) = 2.03⋅10-10 m2/s, c(1) = 0.1 M NaCl, c(2) = 0.001 M NaCl.
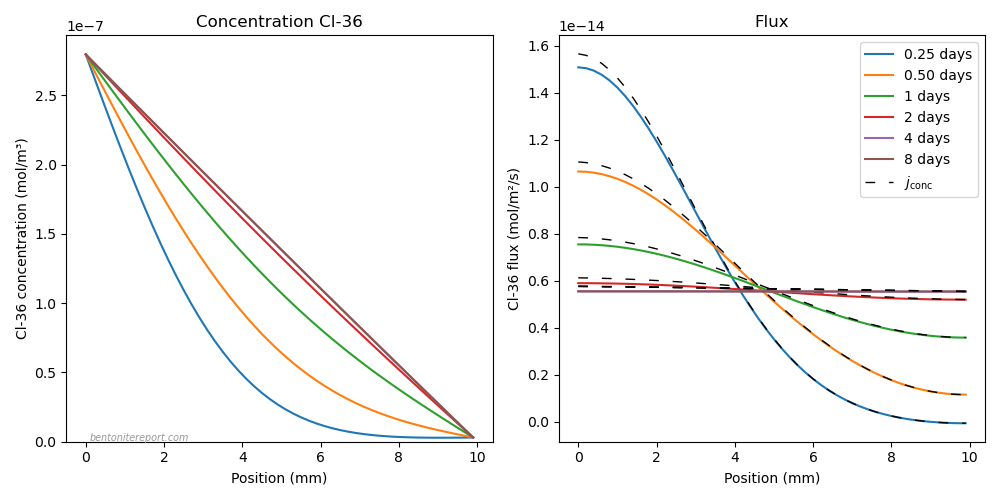
For “example 2”, the concentration and flux evolution for the tracers looks like this
Model parameters: \(c_\mathrm{IL}\)= 0.33 M, \(D_\mathrm{Na}^\mathrm{clay}\) = 1.33⋅10-10 m2/s, \(D_\mathrm{Cl}^\mathrm{clay}\) = 2.03⋅10-10 m2/s, c(1) = 0.1 M NaCl, c(2) = 0.001 M NaCl. c(1)tracers = 10-9 M, c(2)tracers = 10-11 M
In this example, the steady-state flux of both the chloride and the sodium tracer mimics the flux of the main electrolyte; the steady-state flux is the same for both species (5.6⋅10-15 mol/m2/s) and a factor 108 smaller than for the main electrolyte (5.6⋅10-7 mol/m2/s). As the clay incorporates much more 22Na than 36Cl in the transient stage, the initial fluxes are considerably larger for 22Na. The 36Cl flux is in essence completely “Fickian”, i.e. differences between total fluxes and the concentration gradient contributions are negligible. The steady-state 22Na flux, in contrast, has a non-negligible contribution from electromigration, in the same proportion as the stable sodium (this is not resolved in the figure above, but looks qualitatively the same as for the stable sodium, displayed below). These results are completely in line with what we earlier have concluded for the flux of these tracers.
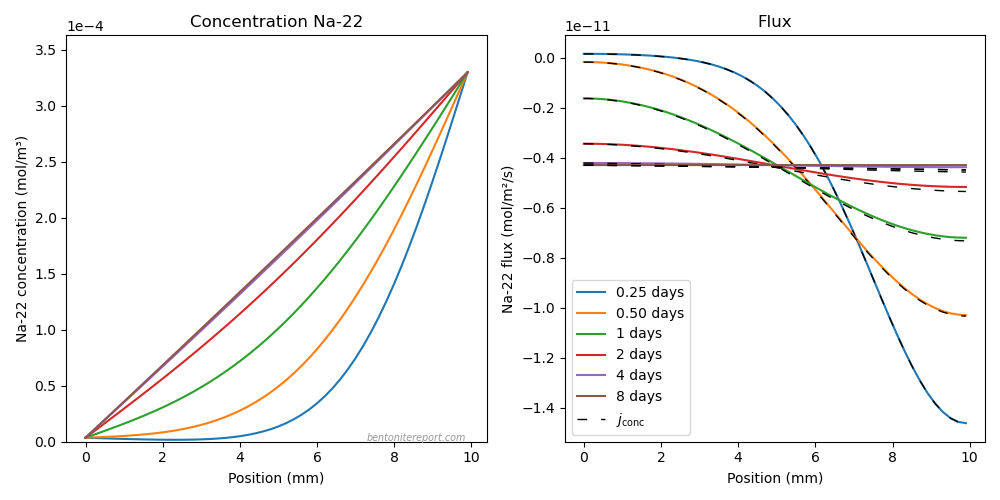
For “example 3”, the tracer concentration and flux evolution looks like this
Model parameters: \(c_\mathrm{IL}\) = 0.33 M, \(D_\mathrm{Na}^\mathrm{clay}\) = 1.33⋅10-10 m2/s, \(D_\mathrm{Cl}^\mathrm{clay}\) = 2.03⋅10-10 m2/s, c(1) = 0.1 M NaCl, c(2) = 0.001 M NaCl. c(1)tracers = 10-9 M, c(2)tracers = 10-9 M
The evolution for 36Cl is basically the same as for “example 2”. This is expected, as the right hand side boundary conditions set the clay concentration at this interface to effectively zero in both cases. The behavior of 22Na, on the other hand, is very different as compared with “example 2”. Now a huge 22Na concentration (relatively speaking) is established at the right hand side boundary, which results in a flux in the opposite direction with much larger magnitude (the steady-state flux is -4.3⋅10-12 mol/m2/s). This illustrates the mechanism behind the seeming uphilldiffusion effect. Note that the 22Na flux in this case is essentially completely “Fickian” during the entire simulation.
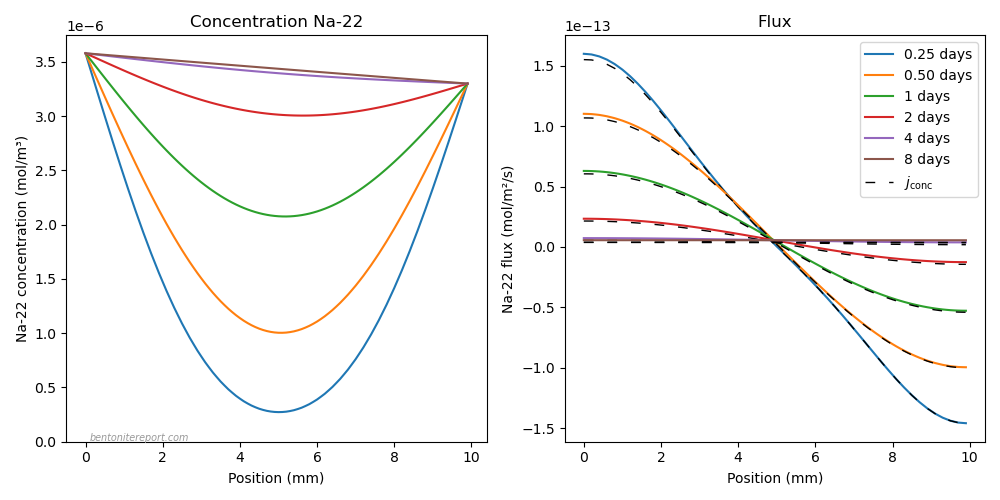
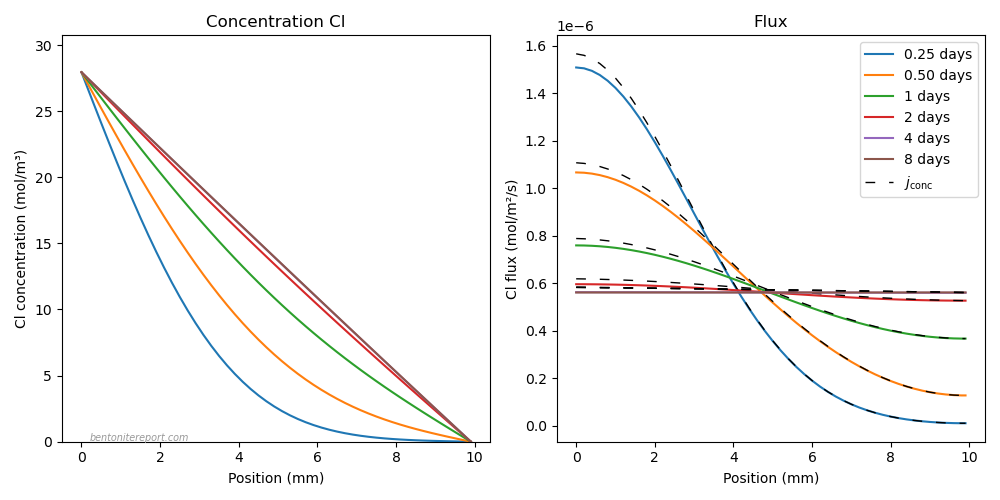
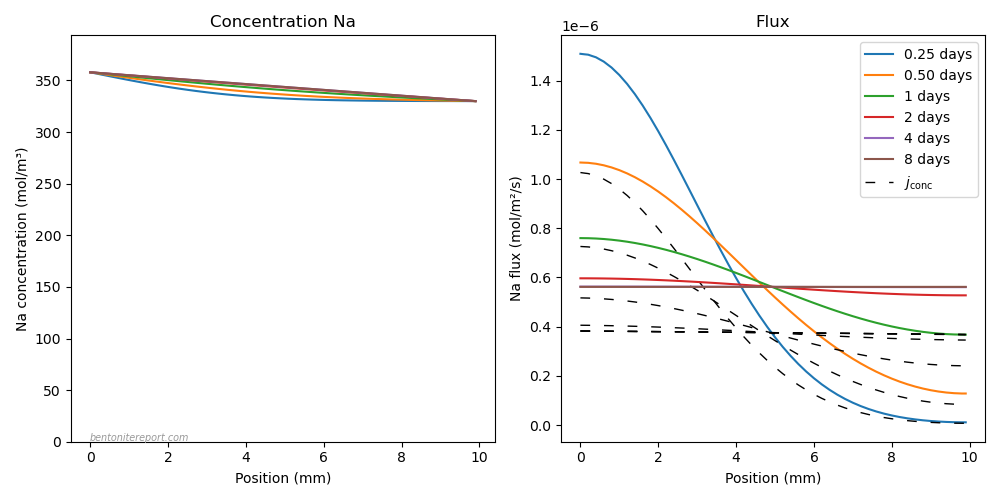
The only significant effect of electromigration in this model is, as we have concluded previously, for the transport of the main electrolyte sodium (or the corresponding 22Na transport in “example 2”, which mimics the main sodium flux). The concentration and flux evolution for main chloride and sodium looks like this (this is the same in both examples).
Model parameters: \(c_\mathrm{IL}\) = 0.33 M, \(D_\mathrm{Na}^\mathrm{clay}\) = 1.33⋅10-10 m2/s, \(D_\mathrm{Cl}^\mathrm{clay}\) = 2.03⋅10-10 m2/s, c(1) = 0.1 M NaCl, c(2) = 0.001 M NaCl.
The total flux are the same for chloride and sodium at all times, as a consequence of the requirement of charge neutrality.4 To achieve this, the sodium flux is “boosted” by the induced electric field (electromigration). The effect is mainly noticeable on the cation, because it has a much larger concentration.
Footnotes
[1] \(\tilde{\psi}\) is normalized by the thermal voltage as \(\tilde{\psi} = \psi/V_T\), where \(V_T = RT/F\). Here we adopt \(V_T\) = 25.7 mV.
[2] The corresponding description for Fickian diffusion in the homogeneous mixture model is found here.
[3] The concentration gradient contributions to the fluxes (\(j_\mathrm{conc}\)) in the diagrams presented here are not color coded (but black), in order for them to be visible also when \(j \approx j_\mathrm{conc}\). I hope it is still reasonably easy to understand which flux profile corresponds to which time.
[4] Strictly, the main chloride and sodium fluxes differ ever so slightly in “example 3”, because there is also an 22Na/stable Na exchange process ongoing. But this difference is not noticeable for the main ions, as the reservoir tracer concentrations are a factor 106 — 108 smaller.
Let’s begin by reminding ourselves where we are in the “story” of the article. TS15 begun by vaguely suggesting that diffusion in bentonite1 cannot be modeled by assuming “Fickian” diffusion. The argument, however, is based on the inability of the traditional diffusion-sorption model to be reasonably fitted to ion through-diffusion data. This inability is not primarily due to assuming Fickian diffusion, but due to assuming immobility of the “sorbed” ions and due to assuming that the pore volume contains nothing but bulk water.
After being promised to be presented “fundamental properties” of bentonite that apparently will demonstrate the problem with assuming “Fickian” diffusion, we were insteadintroduced to a large set of modeling concepts. As a description of actual compacted bentonite, these concepts are mostly irrelevant (e.g. the Gouy-Chapman model), unjustified (e.g. Stern layers on basal surfaces), or simply fantasies (e.g. the idea of “stacks”, or the idea of Donnan equilibrium on the microscopic scale).
Finally, we were presented the Nernst-Planck approach to ion diffusion, which indeed goes beyond a Fickian description. The presented model, however, which includes “bulk water”, “diffuse layer” and “interlayer” domains assumed to be in equilibrium locally, is not suitable for compacted bentonite. Furthermore, no description whatsoever is presented for how this equilibrium is supposed to be maintained. The model is therefore incapable, even principally, to account for swelling — the most profound feature of bentonite in contact with an aqueous solution. Note also that discrimination between “diffuse layer” and “interlayer” domains is based on the pure fantasy of “stacks” as fundamental structural units.
We also showed that the presented Nernst-Planck framework was incorrectly derived, and after having sorted out how it can be reinterpreted,2 we noted that the suggested “concentration” and “diffusion potential” contributions to the diffuse layer flux where completely unfounded.
“From diffusive flux to diffusive transport equations”
In this section I expected to be given an overview of how all the previously introduced model components are supposed to fit together in an overall description for bentonite. With so many parts, there are a large number of quantities to consider. Apart from all the species concentrations in the different “porosity domains”, TS15 have also introduced e.g. electrostatic potentials (eqs. 17, 18, 21, 22, 39, 48, 56 in TS15), adsorption sites on “basal surfaces” (eq. 19 in TS15), “exchanger” sites (eq. 24 in TS15), “layer edge amphoteric” sites (eq. 28 in TS15), models for activity coefficients and their spatial derivatives (eqs. 58, 60, 64 in TS15), and entertained the possibility of having varying sizes of the “porosity domains” (although no constitutive relations has been stated).
Instead, the only thing related to a model description here is a restatement of the mass conservation law. This was already presented in eq. 4 in TS15, but is here incorrectly called the “diffusion equation” (TS15 eq. 65), and stated with an incorrect sign (TS15 eqs. 65, 66 and 68). TS15 express it in its final form as (I have here corrected the sign error, and suppressed an index \(i\))
where \(\phi_\mathrm{bulk}\), \(\phi_\mathrm{DL}\) and \(\phi_\mathrm{exch}\) denotes the bulk water, “diffuse layer” and “interlayer” porosites, respectively; \(c_\mathrm{bulk}\), \(c_\mathrm{DL}\), \(c_\mathrm{exch}\) are the concentrations of the considered species in these “porosity domains”; and \(j_\mathrm{bulk}\), \(j_\mathrm{DL}\), and \(j_\mathrm{exch}\) are the corresponding domain fluxes (presumably given from the previously presented Nernst-Planck framework?).
That’s it.
As the authors stress the importance of adsorption processes throughout the article (even on montmorillonite basal surfaces), I find it very odd to here present an equation that completely omits them. Somewhat ironically, adsorption was included previously, in the section titled “Diffusion basics” (eq. 6 — 8 in TS15). Instead, the above equation (eq. 1) implies mechanisms that may alter the sizes of the “porosity domains” (i.e. \(d\phi/dt \neq 0\), \( d\phi/dx \neq 0 \)). But the article has not presented any constitutive relations for such mechanisms! This whole model presentation feels like having been handed a heap of irregular puzzle pieces with a promise of a grand result, while the reference photo belongs to a different puzzle.
Although eq. 1 provides little concrete information on the actual model, it still manages to contain a fatal flaw: the three domain fluxes are simply added to form some sort of a “total” flux. But this is not how diffusive fluxes work! The first real post I wrote on this blog covered this issue. There we noticed that domain fluxes are added in essentially all publications where multi-porous models are applied to bentonite. To see that separate domain fluxes are not additive should be as simple as considering some specific domain configuration. A trivial example is two domain types connected in series
It is easy to see that the resulting flux in this case cannot be described as a sum of independent domain contributions (e.g, the flux vanishes if \(D_1 = 0\), independent on the value of \(D_2\)). More generally, we may instead refer to one of the favorite references on diffusion in the bentonite research field: Crank (1975) has an entire chapter on this topic (“Diffusion in heterogeneous media”).
It really feels silly to point out, but the knowledge that domain diffusivites generally don’t add was established long before any numerical reactive transport tools were developed. I have trouble expressing what an incredible own goal this procedure is: even if it made sense to partition compacted bentonite into various “porosity domains”, using eq. 1 will still lead to incorrect inferences.
“Applications”
Rather than providing the reader with an actual model overview, TS15 proceed with presenting three different cases, which were modeled with either “CrunchFlowMC” or “PHREEQC”. These case studies are motivated with that they
[…] are used to illustrate the importance of considering coupled diffusion/surface reaction effects in order to understand and to predict migration processes and associated parameters in charged porous media, especially clays.
Let’s keep this formulation in mind as we go through the cases. (Spoiler: we will not be illustrated the importance of much, nor will we be able to understand or predict anything.)
This is a diffusion and “sorption” study of cesium, sodium and iodide in pure Na-montmorillonite. Tracer through-diffusion has been performed for each of these ions in samples of 0.8 g/cm3, at three different background concentrations (NaCl): 0.01 M, 0.1 M, and 0.5 M.
This is the “uphill” diffusion study. It presents the result of two through diffusion tests in pure Na-montmorillonite (1.9 g/cm3), where a Na tracer is observed to migrate from a 0.1 M NaClO4 reservoir to a 0.5 M NaClO4 reservoir, although the tracer concentration in the 0.1 M reservoir is lower than in the 0.5 M reservoir.
This paper presents the results of performing Ca-to-Na ion exchange on initially pure Ca-vermiculite (1.8 g/cm3). Disc-shaped single crystal vermiculite samples were immersed in initially pure NaCl solutions, and the evolution of calcium concentration in these solutions was monitored. The samples were prepared in such a way that out-diffusion only occurred at the outer edge of the vermiculite discs. Tertre et al. (2015) present four such tests, performed with NaCl concentrations 0.003 M, 0.05 M, 0.1 M, and 1.0 M. In addition, three tests were performed with NaCl concentration 0.05 M, which were terminated after 1.94, 7.01, and 18.06 days, respectively. Subsequently, the radial distribution of calcium was measured in these samples, giving “snapshots” of the internal Ca profile.
A common theme of these studies is that they primarily demonstrate effects of exchangeable cations being mobile. Tachi and Yotsuji (2014) clearly show the commonly observed effect of cation “effective diffusivity” becoming increasingly large as the background concentration is lowered, which has been unambiguously demonstrated to be an effect of “interlayer diffusion being the dominant pathway”. Glaus et al. (2013) is a proof-of-concept demonstration of a consequence of the exchangeable cations dominating mass transfer, as I discussed in some detail here. Tertre et al. (2015) assert from the start that “For this type of model system, the interlayer region is the only pore space that contributes to the overall diffusion process […]”.
Some sort of quality analysis of the data being modeled, e.g. a separation of signal and noise. It should also be motivated why certain data is modeled while other is not.
A summary that relates the model results to actual physical processes, and discusses possible implications.
Tachi and Yotsuji (2014)
Without commenting on it, TS15 only model results obtained at background concentration 0.1 M and only in samples of length 10 mm. A reason for this may be that Tachi and Yotsuji (2014) only present reservoir concentration evolutions and final state concentration profiles for all ions at these conditions. However, TS15 anyway disregard the final state profiles and only compare their model with the concentration evolutions. Furthermore, for cesium, the original article contains plots also for background concentrations 0.01 M and 0.5 M. Tachi and Yotsuji (2014) actually present fitted model parameters for all tests. If TS15 were serious about exploring this type of diffusion they could consequently have made a systematic comparison rather than just fitting a model to some rather arbitrarily chosen test results. This is particularly remarkable, as the most prominent feature of tests like this — that the “effective diffusion coefficient” for cations increases essentially without bound with decreasing background concentration, while the same quantity for anions approaches zero — is not really explored, when they only model results from a single background concentration.
It is also clear that TS15 show no interest in the quality of the data. Looking at Tachi and Yotsuji (2014) in a bit more detail, the data has some flaws and inconsistencies. For example, for all background concentrations, Tachi and Yotsuji (2014) report two quite different values of the “anion accessible porosity”,3 depending on whether the sample is 5 or 10 mm long. (TS15 “conveniently” choose the value 0.421 and ignore the value 0.635.) A further inspection also indicates that filter restriction has influenced some of the data, although they have attempted to eliminate such artifacts.4
TS15 don’t clearly state what model components they include, and it is essentially impossible to figure out the exact model used for this exercise. Half way through the section, the model is referred to as the “bulk + diffuse layer water diffusion model”, so it is quite safe to state that they have only included a bulk water and a “diffuse layer” domain. As the description in TS15 is fundamentally based on the (flawed) idea that we can distinguish between “external” and “internal” surfaces in compacted bentonite, it is quite remarkable that the “interlayer” domain is here completely left out. To me, this suggests that TS15 are not themselves especially serious about differing between “diffuse layers” and “interlayers” (after all, this difference is a fantasy, as I have argued throughout the review). This sloppy approach to modeling is, in my opinion, a recipe for ending up with an overparameterized mess. It is also worth noting that TS15 simply fix the size of the the bulk water domain by requiring the model to be “in close agreement” with measured “anion accessible porosity”.5 Thus, after 34 pages of text containing a myriad of mathematical expressions and a specific “consideration of electrostatic effects,” we ultimately just end up with knob-twiddling.
Speaking of electrostatic effects, TS15 don’t comment how aspects of multi-component diffusion influence the results. It is not even clear that the Nernst-Planck framework has been used in this particular modeling. For example, only a single set of fitted tracer diffusion coefficients are reported6 (and no transport parameters are reported for the main electrolyte). Are these supposed to relate to the bulk or the “diffuse layer” domain? I find this lack of discussion quite remarkable, given the emphasis in the rest of the article on the idea that ion diffusion in bentonite is supposed to be fundamentally “non-Fickian”.
TS15 do not even manage to clearly communicate the principal reason as for why the model “makes it possible to calculate the diffusive flux of neutral species, anions and cations with the same conceptual model and with physically feasible parameters.”7 This reason is — of course — that the model allows for exchangeable ions to move and that it provides a mechanism (Donnan equilibrium) for altering the relative ion concentrations in the “diffuse layer” domain. In fact, a more reasonable analysis demonstrates that tracer through-diffusion in bentonite can be satisfactory modeled using a single homogeneous domain for the clay. A homogeneous description is also compatible with basically all other experimental findings in compacted bentonite, as we have discussed massively on the blog.
In contrast, TS15 assume, not only that the montmorillonite in this particular test contains 50% bulk water, but also the existence of a Stern Layer — with corresponding ion immobilization — on montmorillonite basal surfaces. As neither of these assumptions are justified or reasonable, it is clear that the resulting model is overparameterized. Any successful fit thus provides little real information or insight, which also becomes almost comically clear when it is concluded
Results are plotted in Figure 19. The tortuosity values follow the order \(\tau_\mathrm{I^-} < \tau_\mathrm{HTO} < \tau_\mathrm{Na^+} < \tau_\mathrm{Cs^+}\), indicating that the tortuous diffusion pathways are not the same for all of these species. Or alternatively, that the adsorbed species in the Stern layer, considered in the calculations as immobile, are in fact mobile.
The result of the modeling exercise indicates either one thing, or a completely different thing!
Note that TS15 here put forward the bizarre notion that each separate species diffuse in different pore structures, with different “tortuosities”. They apparently don’t dismiss the untenable idea that cesium (with fitted “tortuosity” 0.136) for some reason diffuses though a much less meandering pore network than water (with “tortuosity” 0.047). In reality, the much larger “tortuosity” (which here means a less tortuous network…) for cesium as compared with water indicates that the model underestimates the amount of mobile cesium in the “diffuse layer” and compensates for this by giving it a too large diffusivity. This is signature behavior of an overparameterized model.
How this treatment and way of reasoning is supposed to “illustrate the importance of considering coupled diffusion/surface reaction effects” should be a mystery to any reasonable reader.
Glaus et al. (2013)
Things get even worse, in my opinion, as TS15 next present a modeling of the “uphill” test. Providing a solid explanation for this test could have been a major contribution to the article, as it earlier has claimed that the “uphill” effect is an example of “non-Fickian diffusion processes” and has implied that the effect is caused by non-trival electrostatic couplings. But TS15 seem genuinely uninterested in analyzing how the “uphill” effect arises in their model. Again, the adopted model is not presented in full, and focus is mainly on irrelevant details, such as filter diffusivity.8 The only attempt at describing a mechanism for “uphill” diffusion is half a sentence that says
[…] the macroscopic driving force of the diffusion is the diffusion potential in the diffuse layer, as shown in Figure 12.
But “Figure 12” shows results of “example 3”, from the section on the Nernst-Planck framework (discussed here), rather than results from the actual “uphill” test modeling. The burden of connecting “Figure 12” with the actual “uphill” test is completely left to the reader. I find this very peculiar for several reasons. To begin with, as TS15 evidently have modeled the actual “uphill” test, why don’t they refer to and show the “diffusion potential in the diffuse layer” in the actual model? And although “Figure 12” shows an “uphill” cation flux, the Nernst-Planck transport equations are not solved in “example 3”; in these examples, flux contributions are calculated from imposed linear concentration profiles in a bulk water domain. But since the “uphill” test here has been modeled with a tool that apparently solves the Nernst-Planck transport equations, I find it incomprehensible that these results are not reported. It is also worth noting that TS15 don’t compare their model with the final-state concentration profiles, even though these are readily available in the original publication.
The earlier examples, moreover, assume a density of approximately 1.0 g/cm3 (porosity 0.64), and a bulk water domain that dominates the pore volume. The “uphill” test, in contrast, was conducted at nominal density 1.9 g/cm3 (porosity \(\sim\) 0.3). TS15 themselves explicitly comment that “[a]t this high degree of compaction, the actual presence of bulk water is not certain”.9 Other conditions also differ strongly between these systems. The “uphill” test was performed with NaClO4 as main electrolyte, while “example 3” uses NaCl. “Example 3” has boundary concentrations 0.1 M and 0.001 M for the main electrolyte, while the actual “uphill” test, in contrast, has 0.5 M and 0.1 M.
Disregarding these glaring differences between the actual “uphill” test and “example 3”, the explanation given by TS15 in the above quotation is that the cation tracer ions move “uphill” due to “the diffusion potential in the diffuse layer”. In an earlier part of this review we demonstrated that the way TS15 define the “diffusion potential” contribution to the flux is obvious nonsense, based on misunderstanding the electric potential. To remind ourselves, in “example 3” TS15 make the absurd claim that a tracer concentration profile in the “diffuse layer” that looks like this (this is part of “Figure 12”)
makes NO “concentration gradient” contribution to the flux (because TS15 define such contributions exclusively in terms of bulk water concentrations). Instead, the diffuse layer cation tracer flux in “example 3” is supposed to be driven by an electric field (the “diffusion potential” contribution) in the wrong direction.
We have shown that these preposterous definitions of flux contributions can be corrected, with the correction term involving the gradient of the Donnan potential.10 With reasonable flux definitions, we demonstrated that the “uphill” flux in “example 3” is essentially completely governed by the contribution from the “diffuse layer” concentration gradient (i.e. that the cation tracer flux is “Fickian”). Recently on the blog, we have also calculated the electrostatic potential within the homogeneous mixture model and specifically demonstrated that effects of electromigration on the tracer flux is negligible in the “uphill” test.
Rather than identifying any of the above points, TS15 instead spend a large part of this section discussing that several models, which “are totally different from a conceptual view”, can be fitted to the “uphill” diffusion test data. But all models that can be fitted to the “uphill” test data have this ability because they (i) assume mobility of exchangeable ions and (ii) provide a mechanism for ion equilibrium with bulk water. From the point of view of fitting the data, it is secondary if a model treats the ion equilibrium via “exchange sorption” or Donnan equilibrium,11 or if the compartment containing the exchangeable ions is referred to as “interlayer”, “diffuse layer” or interlayer. That TS15 choose to emphasize that their model fits “equally good”, while it contains several additional and unjustified assumptions, simply indicates that they don’t consider the dangers with overparameterization.
The presentation of the modeling of the “uphill” test in TS15 does not “illustrate the importance of considering coupled diffusion/surface reaction effects in order to understand and to predict migration processes…”. Rather, it obscures process understanding.
Update (261006):The “uphill” test is modeled with the Nernst-Planck framework and the homogeneous mixture modelhere.
Tertre et al. (2015)
As with the previous cases, the description of the modeling of Tertre et al. (2015) lacks sufficient detail, preventing the reader from determining exactly which model was used. Actually, the only model information given is that the data was “reinterpreted using the interlayer diffusion option of PHREEQC”. This must obviously mean that an “interlayer” domain is included in the model, in contrast to the previous cases. But are other domains included? (we have previously been informed, in passing, that “PHREEQC” requires a bulk water domain).
Furthermore, it is not clear if a Stern layer is included in this model, i.e. if some of the exchangeable ions are assumed immobilized, as in the previous cases. As not a word is spoken about this, we reasonably must assume that no Stern layer is included. If this is the case, the inclusion of an immobilization mechanism when modeling Tachi and Yotsuji (2014) and Glaus et al. (2013) becomes even more conspicuous. In fact, TS15 state about Tertre et al. (2015)
This experiment showed unambiguously that interlayer diffusion exists and it made it possible also to quantify the diffusion coefficient for \(Ca^{2+}\) in the interlayer.
I find this statement remarkable for a couple of reasons. As TS15 don’t say anything similar when discussing the other studies, I here get the impression that they haven’t comprehended that “interlayer diffusion” has been unambiguously demonstrated much earlier than in Tertre et al. (2015). Also, if TS15 now acknowledge that interlayer diffusion “exists” in bi-hydrated vermiculite, how come that they don’t assume this to be the primary mode of transport in the other models, but instead add an immobilization mechanism? Are ions supposed to become less mobile in less dense systems with lower layer charge density? We should also remember that earlier in the article, TS15 describe the “interlayer” as pure Stern layers. So, how is it, are Stern/interlayer ions supposed to be mobile or not? Again, I think this relaxed attitude towards when and how various domains are included, and how they are treated, demonstrates that TS15 themselves don’t take the concepts of “diffuse layer” and “interlayer” seriously.12
Furthermore, the “reinterpretation” of this experiment is entirely untenable. Tertre et al. (2015) put a lot of effort in demonstrating (using “Brownian dynamics simulations”) that the observed diffusion is strongly influenced by processes occurring at the interface between the sample and the external solution. This makes sense, as we may expect huge concentration gradients within the clay when two different cations are involved. Such gradients are difficult to maintain, and the resulting flux can instead be expected to be controlled by interface transfer resistance. This is similar to how filters limit the flux in cation through-diffusion tests at low ionic strength. In the “reinterpretation”, TS15 completely neglect any possible interface transfer resistance, claiming
It was possible to reproduce the data with the same level of quality as the Brownian dynamics calculations by considering an interlayer \(Ca^{2+}\) diffusion coefficient value of \(0.8\times 10^{-11}\) \(\mathrm{m}^2\cdot \mathrm{s}^{-1}\), together with an interlayer \(Na^{+}\) diffusion coefficient of \(4\times 10^{-11}\) \(\mathrm{m}^2\cdot \mathrm{s}^{-1}\), \(1\times 10^{-11}\) \(\mathrm{m}^2\cdot \mathrm{s}^{-1}\), \(0.1\times 10^{-11}\) \(\mathrm{m}^2\cdot \mathrm{s}^{-1}\), and \(0.1\times 10^{-11}\) \(\mathrm{m}^2\cdot \mathrm{s}^{-1}\), for the experiments at NaCl concentration of 1, 0.1, 0.05 and 0.003 \(\mathrm{mol}\cdot \mathrm{L}^{-1}\) respectively. The decrease of the \(Na^{+}\) interlayer diffusion coefficient with NaCl concentration is in agreement with MD results from Tertre et al. (2015), which show a decrease of its value with an increase of the \(Ca^{2+}\)/\(Na^{+}\) occupancy ratio in the interlayer (Fig. 21c).
The last sentence is false. Tertre et al. (2015) indeed report a decrease of the sodium self-diffusion coefficient, but this decrease is lower than a factor 3.13 In the model of TS15, in contrast, this decrease is a factor 40 (!), which certainly cannot be considered “in agreement”. In addition, experimentally, Kozaki et al.(2005) observed no systematic decrease of Na diffusivity with increasing Ca fraction, in bi-hydrated montmorillonite; if anything, they observe a slight decrease (25 — 60%) of Na diffusivity only in pure Ca-montmorillonite. TS15 anyway goes so far as to conclude this part by stating
Consequently, the apparent decrease in \(Ca^{2+}\) interlayer diffusion coefficient can also be interpreted as a result of the decrease of the \(Na^{+}\) interlayer diffusion coefficient that arises from the coupling between the diffusion of these two species through the diffusion potential term in Equation (57).
Let’s unpack the serious error of reasoning on display here: rather than taking into account interface transfer resistance — a process that we have every reason to believe is active, and that is expected to produce the observed behavior — the observed behavior is supposed to be caused by an unjustified 40-fold decrease of the sodium self-diffusivity, as the external concentration decreases. A mobility decrease that do not even correlate directly with the calcium content of the clay! Note also that TS15 state that the decrease in sodium diffusivity is caused by electromigration (“the diffusion potential term in Equation (57)”). But, as far as I understand, these sodium diffusivities were just put in by hand! (See previous quotation.) It is completely unreasonable that TS15 are here allowed to provide this “explanation” without showing a single result from the actual modeling of how this “diffusion potential term” is supposed to change under different conditions.14
It feels silly to point out that this modeling does not in any way “illustrate the importance of considering coupled diffusion/surface reaction effects in order to understand and to predict migration processes…”. Instead, it is a horrific example of “modeling” as a game, where the only aim is to be able to fit a model to some arbitrary data.
The overall impression of these modeling cases is that TS15 do not seem interested in understanding or communicating how compacted bentonite really works: models are not fully described, and components seem included or disregarded pretty much on a whim; no attention is paid to the quality of the data that is modeled, nor to how data is chosen; and discussions are completely absent regarding how the modeling can aid process understanding. I also find it curious how little these modeling cases connect with the main topics discussed in the rest of the article. For example, very little is said about the impact of multi-component diffusion, even though “non-Fickian” diffusion is a a major theme of the article (it is often not even clear when and how multi-component diffusion has been implemented). Likewise, although sorption processes are included in some of the model cases, we are not told anything about their significance. And not a word is spoken about evolving volumes of the various “porosity domains”, even though such processes seem to be taken seriously in other parts of the article.
“Summary and perspective”
We have finally arrived at the last page of the article. The first paragraph of this section mainly concerns the shortcomings of the traditional diffusion-sorption model (“the classical Fickian diffusion model”)
In this context, the classical Fickian diffusion model applied in the framework of a single pore diffusion model with linear adsorption processes (\(K_D\) model) appears to be inappropriate for describing diffusion data without making less than satisfying modeling assumptions, such as i) different definitions of the porosity as a function of the nature of the tracer of interest (see anion accessible porosity) and as a function of the conditions (anion accessible porosity change with time; ii) change of the adsorption parameters from batch to diffusion experiment; and iii) physically unrealistic pore diffusion coefficients (or tortuosity values).
I agree with that the traditional diffusion-sorption model is not suitable for bentonite. But this has been known for a long time, as I have argued from the beginning of this review. That TS15 use this model as a starting point — and argues for modifying it rather than dismissing it — is in my head a recipe for disaster.
I want to emphasize that all empirical evidence points to that compacted water saturated bentonite is a “single pore” material. The main problem with the traditional diffusion-sorption model is not that it assumes a “single pore”, but that this “single pore” is assumed to contain bulk water. Letting go of the bulk water allows for a satisfying bentonite model using a single pore type. I’ve elaborated on these points here and here.
This first paragraph also says
Diffusion processes through clay materials is the result of a complex interplay of transport and non-linear adsorption processes under the influence of electrostatic fields.
This formulation must be interpreted as summarizing what has been covered in the article as a whole. But I don’t agree with this description, and throughout this review we have exposed flaws and shortcomings in much of the presented material. Of course, ion diffusion is generally influenced by electric fields, since the diffusing entities carry charge. But we have shown that the definition of a “diffusion potential” made by TS15 makes no sense and is based on misunderstanding how electric potentials function. With corrected definitions we have further shown that processes described by TS15 as being due to a “diffusion potential” is actually mainly driven by concentration gradients, i.e. the diffusion processes are “Fickian”. I see it as a major problem that many researchers don’t recognize diffusion in compacted bentonite as simpler than usually described.
This sentence also claims that “non-linear adsorption processes” are involved generally in ion diffusion in bentonite. But this is not the case! The ions that the article mostly focuses on — e.g. sodium, calcium, chloride and iodide — don’t show any signs of adsorption15 at all, let alone “non-linear” adsorption.
The second paragraph of this section is fully devoted to “non-linearity of adsorption processes for strongly adsorbed species”, with particular focus on cesium. I find this peculiar, as non-linear sorption is only brought up in a single small part in the article, and as nothing is said about non-linearity in the the reported “application” that involves cesium (Tachi and Yotsuji, 2014)
The third paragraph begins
Recent developments of selected RT [reactive transport] codes that can handle diffusion processes in diffuse layer and interlayer porosities made it possible to model the diffusion data of neutral, anionic and cationic species within the same conceptual framework.
Here is presumed that partitioning of the pore space into “diffuse layer” and “interlayer” domains makes sense. But we have thoroughly discussed that this whole notion — which is based on the fantasy concept of “stacks” as fundamental structural units of compacted bentonite — makes essentially no sense. And, as we noted earlier, TS15 themselves employ these “porosity domains” for convenience, rather than from being constrained by empirical evidence.
Moreover, it is not the ability to define both “diffuse layer” and “interlayer” that makes it possible to treat any type of species “within the same conceptual framework”16 Rather, a coherent model for compacted bentonite is made possible by realizing that exchangeable ions are mobile and by treating the corresponding ion equilibrium when required. This is (and has been) most adequately made in a homogeneous model.
The third paragraph continues
Also, the contribution of the diffuse layer and/or the interlayer to the overall diffusion of ions makes it possible to explain the origin of the apparent acceleration of cation diffusion as compared to water, which otherwise would require unrealistic tortuosity values for cations in classical Fickian diffusion models.
It is not true generally that cation diffusion is “accelerated”. Speaking of “accelerated” cation fluxes is mostly relevant for steady-state in conventional through-diffusion tests at low background concentration. Writing like this makes me suspect that TS15 have not identified this distinction, especially since they do not recognize the importance of interface equilibrium. As I discussed in the first part of the review, e.g. sodium and chloride show almost identical diffusive behavior in bentonite. The additional effects observed in through-diffusion is completely due to Donnan equilibrium at the interfaces to the external solutions, as has been explained in detail long before the publication of the present article. Again, TS15 contrast with “classical Fickian diffusion models” — but it has been fully clear from the start that the traditional diffusion-sorption model does not apply to compacted bentonite. The way TS15 here use the phrasing “diffuse layer and/or interlayer” again indicates that they don’t take their own concepts seriously. The third paragraph ends
RT modeling has also helped to improve our understanding of anomalous diffusion behavior such as that of up-hill diffusion.
This is a horrible statement. That they refer to the “uphill” effect as “anomalous” tells me that they have neither understood this experiment, nor how bentonite works.17 And if they by “RT modeling” mean the previously presented modeling of the “uphill” effect, nothing can be further from the truth than claiming that it has improved understanding. I also find it significant that they don’t connect the “uphill” test to the previous statements in the same paragraph; the “uphill” test was designed to demonstrate that cation diffusion is not simply “accelerated” (it is, in a sense, conventional tracer through-diffusion that is “anomalous”).
The fourth paragraph reads
Despite the successes of these new RT modeling approaches, it must be stressed that the model and its parameters derived from diffusion experiments are not always unique. Two examples given above highlight the fact that several different conceptual models can provide equally good fits of the data. As such, the modeling effort is typically under constrained, a fact that explains the multitude of conceptual and numerical models available in the literature that describe the ionic transport properties of clay media (Leroy et al. 2006; Appelo and Wersin 2007; Gonçalvès et al. 2007; Birgersson and Karnland 2009; Gimmi and Kosakowski 2011; Tachi et al. 2014).
I strongly disagree with this whole line of reasoning. The multitude of models is not explained by them being “under constrained”; it is explained by most of them being overparameterized (which is poison).
Serious modeling work requires that all included components be evaluated and justified on physical grounds. I see very little of this in the contemporary bentonite scientific literature. We can use the “uphill” test modeling to illustrate this point. The model of this test in TS15 includes, among other things, a mechanism for sodium adsorption and immobilization on basal surfaces (a Stern layer). The test, however, has already been demonstrated to be well described by models that don’t include such a mechanism. This test can therefore not in itself be used to motivate the inclusion of a Stern layer. On the contrary, if there are no other reasonable arguments for why sodium is supposed to adsorb and become immobilized in montmorillonite, it is bad modeling practice to include a Stern layer component.
To make an absurd analogy: if I, as a modeler, don’t have to motivate the included components, I may claim that bentonite contains invisible unicorns that eat anions (or whatever), as long as my model can be fitted to some arbitrary data. This may sound funny, but at least in my head, contemporary bentonite models are quite full of “unicorns”, e.g.
They include significant amounts of bulk water
They rely on the fantasy concept of stacks
Equilibrium between various domains is maintained without a mechanism
They postulate regions completely devoid of anions
Diffusive fluxes from various domains are added
Simple ions, e.g. sodium, are assumed to bond covalently (or whatever) to montmorillonite basal surfaces
In the last paragraphs of this section, TS15 argue for that more research is needed to overcome the claimed problem of models being “under constrained”, using e.g. molecular dynamics simulations and developing new observation techniques. In contrast, I firmly believe that for modeling activities to be meaningful, researchers instead need to spend much more effort scrutinizing and analyzing their models — and the data they are fitting them to. This is much of what I’m trying to do on this blog.
Final comments
My main interest in doing this review has not been in the specific article that we have dissected, but in how it reflects the state of a broader part of the bentonite research field. After finishing, my initial opinion is strengthen: this research field is in a terrible state. I am now fully convinced that reasonable support is nowhere to be found for several of the concepts frequently adopted in contemporary bentonite descriptions. This applies in particular to the ideas of “stacks” as fundamental structural units, the presence of bulk waterwithin the bentonite, and a general adsorption/immobilization mechanism for ions on montmorillonite basal surfaces (Stern layer). To me, it is also clear that this research field lacks knowledge of how salt exclusion and swelling actually work in compacted bentonite, as well as insight into how central these phenomena are for an adequate description.
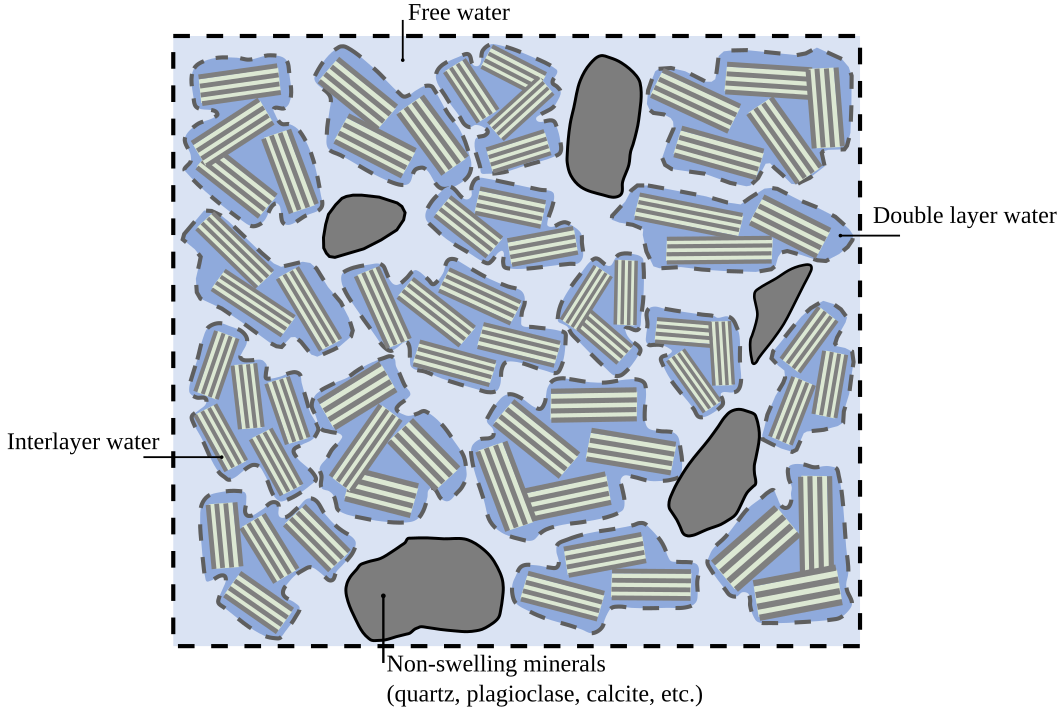
In today’s bentonite scientific literature, you can simply close your eyes and point at random to find articles that peddle much of the type of nonsense listed above. An illustrative (literally!) example of this legacy is found in Chen et al. (2024)
In the past decades, scholars have carried out a lot of research on the pore structure of compacted bentonite, and it is now generally accepted that there are three different types of pores, including inter-aggregate pores, inter-particle pores and interlayer (or inter-laminar) pores (Lloret and Villar, 2007; Wang et al., 2014), as shown in Fig. 1.
[…]
In existing research, water existing inter-aggregate pores, interparticle pores and interlayer pores are usually referred to as free water, double layer water and interlayer water. In general, anions only diffuse in free water because anions are expelled from the interlayer water and double layer water owing to exclusion effect (Glaus et al., 2007; Wu et al., 2020). Adversely [sic] cation diffusion is supposed to occur in all three types of water due to positive adsorption, corresponding to free water diffusion, surface diffusion and interlayer diffusion, respectively (Melkior et al., 2009; Tachi and Yotsuji, 2014; Yang and Wang, 2019)
“Fig. 1” in Chen et al. (2024) looks very similar to this18
The saddest aspect of this quotation is that it is partly correct: “Scholars have carried out a lot of research” for “decades”, and this view is “generally accepted”.
I cannot say that doing this review has answered my question as to why this is the state of the bentonite research field, but maybe a hint is given by the attitude expressed in TS15 towards modeling and model development. I believe this field will have trouble progressing as long as “modeling” is synonymous with getting arbitrary mathematical descriptions to fit arbitrarily chosen data, padded with empty talk.
Footnotes
[1] As I have commented in the earlier parts: TS15 are fond of using the general terms “clays” and “clay minerals”, while it is clear that the publication mainly focus on systems with substantial ion exchange capacity and swelling properties. Here we will continue to use the term “bentonite” for these systems, and ignore the frequent references in TS15 to more general terms.
[2] Such reinterpretation involves the gradient of the bulk water electrical potential — a potential that TS15 put strictly equal to zero. It also involves the gradient of the Donnan potential — a potential that TS15 mistake for the electric potential in the diffuse layer, when it actually describes the potential difference between bulk and diffuse layer.
[3] Tachi and Yotsuji (2014) report “rock capacity factors” for iodide, which erroneously often are interpreted as anion-accessible porosities. TS15, as we have seen, go “all in” on this concept, while simultaneously admitting that it is fictitious. (?!)
[4] We have noted earlier that it seems very difficult to completely eliminate the effects of interface transfer resistance.
[5] Note that the “correct” value is the arbitrarily selected experimental result 0.42, while the other experimental result of 0.64 is ignored. Also remember that the way this parameter varies with ionic strength is not at all explored by TS15, as they only model results from tests with 0.1 M background concentration.
[6] TS15 report “tortuosities”, rather than actual diffusion coefficients. In a bentonite context, the only way such quantities make sense is as a measure of the diffusion coefficients in terms the corresponding bulk water value, as we discuss here.
[7] I object to that the model parameters are “feasible”. It is in my head a completely ludicrous idea that half the pore volume in pure sodium montmorillonite at 0.8 g/cm3 should contain bulk water — the swelling pressure for a 0.1 M NaCl background concentration is expected to be above 0.4 MPa! Also, the idea that the diffusivity of cesium in montmorillonite is much larger than for sodium, iodide or water is completely unreasonable. In fact, TS15 state that “feasible” simply means a cesium diffusivity that is smaller than for pure bulk water. This shows that they are under the impression that “\(D_e\)”, rather than “\(D_a\)”, quantifies the actual diffusivity in the clay. For a further discussion on this, see here.
[8] Including filter diffusivity in this modeling is certainly not irrelevant per se. However, TS15 merely assign a seemingly random value to this quantity and do nothing more about it. All subsequent inferences are thus contingent on this particular value, yet the authors have neither demonstrated if nor how it influences the overall process.
[9] Although I agree with that these systems don’t contain any appreciable amount of bulk water, we note that no reference is given for this claim. I find this annoying, as TS15 simultaneously don’t hesitate to ascribe 50% bulk water to pure Na-montmorillonite at 0.8 g/cm3. As commented earlier, such a system is expected to have swelling pressure above 0.4 MPa.
[10] We still found serious problems with the model formulation, as further discussed in part 5 of the review.
[12] As always, we should also remember that the distinction betweeen “diffuse layers” and “interlayers” (with quotation marks), or similarly, between “inner” and “outer” basal surfaces, is pure fantasy.
[13] It’s not completely straightforward to extract exact MD values from Tertre et al. (2015). But, discussing them in terms of “surface mobility”, they write: “the mobility for Na is variable from approximately 0.04 to approximately 0.015 when the equivalent fraction of Ca in the solid increases from 0 to 1.” This is a decrease by a factor 2.7.
[14] Note, for instance, that sodium is set to be faster than calcium for the cases of 1.0 M and 0.1 M NaCl, while it is set to be slower than calcium for the cases of 0.05 M and 0.003 M NaCl. Is calcium supposed to be able to retard the sodium flux to such an extent that the sodium ion in the end diffuses slower than the calcium ion?! Note also that the sodium mobility in the TS15 model is lowered by an order of magnitude as the external concentration is only lowered from 0.1 M to 0.05 M! (Or, if the corresponding figure rather than the text is correct, the mobility is even lowered by a factor 40 (the later sections of the paper contain an unacceptable amount of typos of this kind).)
[15] Of course, if ion exchange is supposed to be an “adsorption” process, any cation do “adsorb”. But using such a label for a process that actually may enhance the diffusive flux is, in my opinion, a very bad idea.
[16] To me, the “conceptual framework” promoted in TS15 is “add stuff as you please (but keep bulk water at all costs)”.
[17] Calling the “uphill” effect “anomalous” also seems to contradict the previous general statement that “Diffusion processes through clay materials is the result of a complex interplay of transport and non-linear adsorption processes under the influence of electrostatic fields.”
[18] This only represents a part of “Fig. 1” in Chen et al. (2024), which is explicitly stated to represent compacted water saturated bentonite. I will not elaborate on what is wrong with this figure, but here are some questions to ask: 1. Does this figure represent a realistic size distribution of montmorillonite TOT-layers? 2. Does it represent a typical size distribution of accessory minerals? 3. What is the effective montmorillonite dry density of the depicted system? 4. How is swelling pressure maintained? 5. What keeps “particles” and “aggregates” together? 6. How did this illustration pass peer-review?
Lately on the blog we have had a certain focus on the electric potential in compacted bentonite systems. In the ongoing post-publication review of Tournassat and Steefel (2015), for example, we have deep-dived into how electric potentials are treated in multi-porous models. We have also discussed the seeming “uphill” diffusion effect in different contexts. Although this effect is principally explained by the same mechanisms as for traditional cation tracer through-diffusion, we should address the potential difference that is present across the sample in an “uphill” test.
Here is presented a somewhat more general description of the electric potential difference across a bentonite sample, within the homogeneous mixture model. This treatment is essentially equivalent to how this so-called membrane potential is conventionally evaluated within the ion-exchange membrane scientific discipline. As usual on the blog, the message here is that compacted bentonite should not be modeled with various “porosities” and sorption “sites” (for ion exchange). Rather, from the perspective of ion transport,1 compacted bentonite appears to be well described as a conventional charged ion-exchange membrane.2
Multi-component diffusion in the homogeneous mixture model
The external solutions (labeled “1” and “2”) are specified by general species compositions, with the only constraint that they should be electrically neutral. We consequently consider a situation with possible chemical gradients — and corresponding mass transfer — across the bentonite sample, while the hydrostatic pressures in the two reservoirs are assumed equal.3 The values of the electric potential in the external solutions have been labeled \(\psi^{(1)}\) and \(\psi^{(2)}\), respectively.
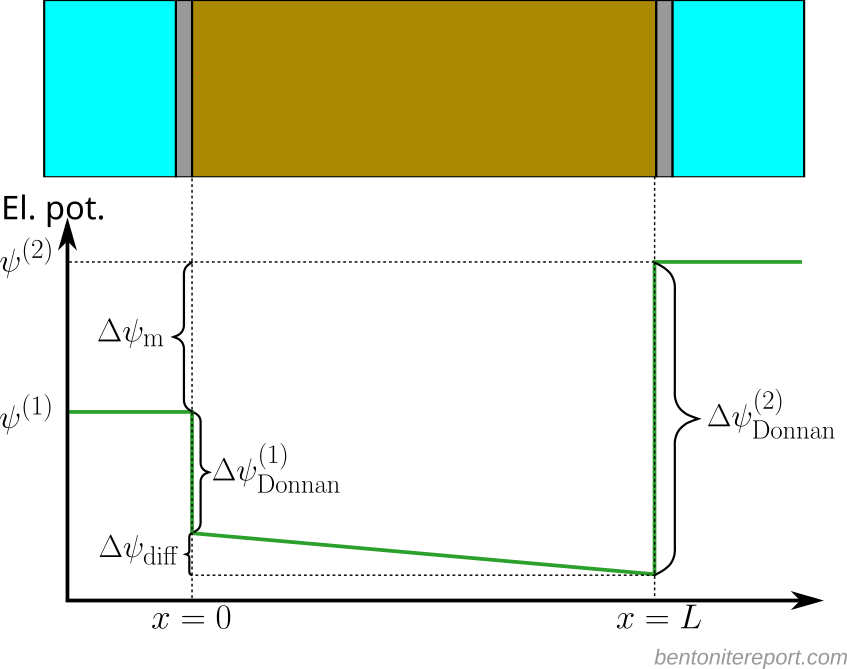
Treating the bentonite as a single homogeneous phase, the electric potential has discontinuities at the external solution/clay interfaces, due to Donnan equilibrium. In addition, the potential may vary within the bentonite, due to diffusion of charged species with different mobilities (electromigration). Schematically, the steady-state electric potential profile can be illustrated like this
Here, \(\Delta \psi_\mathrm{Donnan}^{(1)}\) and \(\Delta \psi_\mathrm{Donnan}^{(2)}\) denote the potential changes due to Donnan equilibrium at the two interfaces.4 \(\Delta \psi_\mathrm{diff}\), which conventionally is named the diffusion potential, denotes the total change in potential across the sample due to electromigration. The membrane potential, \(\Delta \psi_\mathrm{m}\), can be expressed
Given the reservoir concentrations, the Donnan contributions can be calculated using the general framework, if also a value of \(c_\mathrm{IL}\) — which expresses the montmorillonite structural charge as a monovalent interlayer species concentration — is specified for the bentonite component. Such calculations also provide the corresponding sets of internal interface concentrations, i.e. \(\left \{c_{1}^\mathrm{int}(0), c_{3}^\mathrm{int}(0), c_{3}^\mathrm{int}(0), \ldots \right \}\), and \(\left \{c_{1}^\mathrm{int}(L), c_{3}^\mathrm{int}(L), c_{3}^\mathrm{int}(L), \ldots \right \}\), where we denote by \(c_i^\mathrm{int}(x)\) the concentration for species \(i\) at position \(x\) within the clay, and we assume the clay domain located between \(x=0\) and \(x=L\).
The sets of internal interface concentrations, in turn, constitute boundary conditions for solving e.g. the Nernst-Planck equation in the clay domain. Within the Nernst-Planck framework, the gradient of the electrostatic potential in the clay is expressed in terms of species concentrations and diffusivities as
where \(z_i\) and \(D_i\) denote, respectively, the charge number and diffusion coefficient in the clay domain, for species \(i\), and \(V_T = RT/F\) is the thermal voltage (we assume \(V_T\) = 25.7 mV here). From eq. 2 the diffusion potential can be evaluated.
Note that the problem of ending up with a too constrained model when both the Donnan equilibrium and the Nernst-Planck frameworks are invoked does not occur here. This is in contrast to the problems we recently discussed in the model of Tournassat and Steefel (2015). That model requires Donnan equilibrium at each point within the clay, while here it is only imposed at the two interface points.
In the following we will evaluate the membrane potential numerically in some different cases. We start, however, by analyzing the 1:1 system analytically, in order to get a better feel for these processes in compacted bentonite.
1:1 system
The pure 1:1 system has a single type of monovalent cation and a single type of monovalent anion (e.g. Na-montmorillonite in contact with NaCl solutions). The requirement of charge neutrality dictates equal concentrations of cations (\(c_+\)) and anions (\(c_-\)) in each external solution, and we write
where \(c^{(n)}\) is the electrolyte concentration in reservoir \(n\) (\(n\) = 1 or 2).
In this analytical treatment, we assume the often relevant condition of having small external concentrations as compared with the concentration of structural charge in the clay, i.e.
where \(\psi^{\star, (n)}\) is the Donnan potential. In eq. 6 we have ignored the activity coefficient correction factor; contributions to the membrane potentials from activity coefficient differences are expected to be small, and here we will not consider them.5 Note that Donnan potentials are negative, and that \(\Delta \psi_\mathrm{Donnan}^{(1)} = \psi^{\star, (1)}\) and \(\Delta \psi_\mathrm{Donnan}^{(2)} = -\psi^{\star, (2)}\). Utilizing eq. 6, we can express the total Donnan contribution to the membrane potential as6
where we also have continued assuming low external concentrations (eq. 4).
If we impose a constant internal concentration gradient, i.e. \(\nabla c_-^\mathrm{int} (x) = \Delta c_-^\mathrm{int}/L\), where \(\Delta c_-^\mathrm{int} \equiv c_-^\mathrm{int}(L) – c_-^\mathrm{int}(0)\), the diffusion potential can be written
The first term — the ideal Donnan contribution — depends only on the ratio of the concentration in the two reservoirs, and usually dominates over the diffusion potential (second term).
The diffusion potential is expected to be small under most circumstances, as it is seen to explicitly depend on \(1/c_\mathrm{IL}\). This behavior reflects that potential variations within the bentonite are reduced due to the presence of counter-ions (bentonite is a decent conductor). But the factor \(\Delta c_-^\mathrm{int}\) in eq. 11 contributes with an additional factor \(1/c_\mathrm{IL}\) to the diffusion potential, because of Donnan equilibrium (see eq. 6)
Note also that the diffusion potential only depends on the relative values of the diffusivities, and consequently vanishes if there is no difference in mobility between the cation and the anion.
Comparisons with experiment
We can compare the present theory and approximations with measurements reported in Yaroshchuck et al. (2007). In this study, the membrane potential (called the “concentration potential”) was measured across sodium montmorillonite of density 2.0 g/cm3, with one external solution kept at 0.1 M NaCl and the other raised to 0.2 M, 0.3 M, or 0.5 M. The very high density corresponds to a value of \(c_\mathrm{IL} \approx\) 6.0 M,7 and thus eq. 12 applies (see eq. 4).
In order to also investigate the influence of a diffusion potential we should adopt values for the individual diffusivities of chloride and sodium. Experimental tracer diffusion results indicate that these ions have quite similar mobility. E.g., in pure Na-montmorillonite at 1.8 g/cm3 and 298 K, Kozaki et al. (1999, 2001, 2005) report values in the range \(1.5\cdot 10^{-11}\) — \(2.5\cdot 10^{-11}\) m2/s for chloride, and \(2.1\cdot 10^{-11}\) — \(2.4\cdot 10^{-11}\) m2/s for sodium. From these values we can estimate a range for the factor (\(D_-/D_+-1\)) of -0.4 — 0.2. Rather than adopting such a value, however, we here assume the diffusivities to have the corresponding bulk water ratio (1.53), giving \((D_-/D_+ -1)\) = 0.53. This is similar to the treatment in the examples in Tournassat and Steefel (2015), which we have discussedrecently. Theoretical and experimental results are compared in the table below
Evaluations using cIL = 6.0 M, c(2) = 0.1 M, (D–/D+ – 1) = 0.53:
App-
rox-
ima-
tion
Full
Frame-
work
Exp.
\(c^{(1)}\) (M)
\(\psi^{\star,(1)}\) (mV)
\(\psi^{\star,(2)}\) (mV)
\(\Delta \psi_\mathrm{D}\) (mV)
\(\Delta\psi_\mathrm{diff}\) (mV)
\(\psi_\mathrm{m}\) (mV)
\(\psi^{\star,(1)}\) (mV)
\(\psi^{\star,(2)}\) (mV)
\(\Delta \psi_\mathrm{D}\) (mV)
\(\Delta\psi_\mathrm{diff}\) (mV)
\(\psi_\mathrm{m}\) (mV)
\(\psi_\mathrm{m}\) (mV)
0.2
-87.4
-105.2
17.8
-0.011
17.8
-87.4
-105.2
17.8
-0.011
17.8
18
0.3
-77.0
-105.2
28.2
-0.030
28.2
-77.1
-105.2
28.1
-0.030
28.1
28
0.5
-63.9
-105.2
41.4
-0.090
41.3
-64.0
-105.2
41.2
-0.088
41.1
41
This table compares calculations performed using eq. 12 (columns 2 — 6), calculations performed using the full frameworks for Donnan equilibrium and the Nernst-Planck potential (columns 7 — 11), and experimental values (last column). We note that, for this very high value of \(c_\mathrm{IL}\), the calculated Donnan contributions (\(\Delta \psi_\mathrm{D}\)) are essentially identical whether or not the full framework is adopted. We also see that the calculated diffusion potential contributions are completely negligible: the largest value calculated is still below 0.1 mV! Finally, we note excellent agreement between calculated and measured membrane potentials.
The agreement with experiment suggests, not only that the membrane potential is completely determined by Donnan equilibrium in these systems, but that bentonite essentially play by the same rules as conventional ion-exchange membranes.2 In particular, these results8 are even more evidence for that all exchangeable cations are mobile in bentonite.
To explore the NaCl system further, let’s investigate how the results change for lower clay density. In the tables below are shown corresponding results as above, but for \(c_\mathrm{IL}\) = 3.0 M and \(c_\mathrm{IL}\) = 1.5 M, which roughly correspond to densities 1.6 g/cm3 and 1.1 g/cm3, respectively.
Evaluations using cIL = 3.0 M, c(2) = 0.1 M, (D–/D+ – 1) = 0.53:
App-
rox-
ima-
tion
Full
Frame-
work
\(c^{(1)}\) (M)
\(\psi^{\star,(1)}\) (mV)
\(\psi^{\star,(2)}\) (mV)
\(\Delta \psi_\mathrm{D}\) (mV)
\(\Delta\psi_\mathrm{diff}\) (mV)
\(\psi_\mathrm{m}\) (mV)
\(\psi^{\star,(1)}\) (mV)
\(\psi^{\star,(2)}\) (mV)
\(\Delta \psi_\mathrm{D}\) (mV)
\(\Delta\psi_\mathrm{diff}\) (mV)
\(\psi_\mathrm{m}\) (mV)
0.2
-69.6
-87.4
17.8
-0.045
17.8
-69.7
-87.4
17.7
-0.044
17.7
0.3
-59.2
-87.4
28.2
-0.120
28.1
-59.4
-87.4
28.0
-0.116
27.9
0.5
-46.0
-87.4
41.4
-0.361
41.0
-46.7
-87.4
40.7
-0.336
40.4
Evaluations using cIL = 1.5 M, c(2) = 0.1 M, (D–/D+ – 1) = 0.53:
App-
rox-
ima-
tion
Full
Frame-
work
\(c^{(1)}\) (M)
\(\psi^{\star,(1)}\) (mV)
\(\psi^{\star,(2)}\) (mV)
\(\Delta \psi_\mathrm{D}\) (mV)
\(\Delta\psi_\mathrm{diff}\) (mV)
\(\psi_\mathrm{m}\) (mV)
\(\psi^{\star,(1)}\) (mV)
\(\psi^{\star,(2)}\) (mV)
\(\Delta \psi_\mathrm{D}\) (mV)
\(\Delta\psi_\mathrm{diff}\) (mV)
\(\psi_\mathrm{m}\) (mV)
0.2
-51.8
-69.6
17.8
-0.18
17.6
-52.2
-69.7
17.5
-0.170
17.3
0.3
-41.4
-69.6
28.2
-0.48
27.8
-42.3
-69.7
27.4
-0.434
26.9
0.5
-28.2
-69.6
41.4
-1.44
39.9
-30.7
-69.7
39.0
-1.146
37.8
For \(c_\mathrm{IL}\) = 3.0 M, the difference between using the full framework or eq. 12 is seen to still only affect the evaluated membrane potentials on the order of tenths of mV. Only for the higher external concentrations in the system with \(c_\mathrm{IL}\) = 1.5 M are the differences becoming above 1 mV. Similarly, the diffusion potential is seen to be essentially negligible for all these cases.
Further investigation of potentials and fluxes
It should be emphasized that we are not actually solving the full Nernst-Planck transport framework in these examples. Rather, we calculate the corresponding potential difference as we impose a linear internal concentration profile. This is similar to the evaluation made by Tournassat and Steefel (2015) — although they impose concentration profiles in a presumed bulk water phase within the clay. Similar to that publication, we will here visualize the corresponding fluxes in some cases of interest.
The total flux of a charged species in the Nernst-Planck framework can be written as the sum of a “Fickian” contribution (i.e. proportional to the corresponding concentration gradient) and a contribution from electromigration (i.e. flux induced by the presence of an electric field).9
Below are plotted imposed concentration profiles, Nernst-Planck fluxes, and the electric potential profile, for the NaCl-system with \(c_\mathrm{IL}\)=3.0 M, \(c^{(1)}\) = 0.5 M, and \(c^{(2)}\) = 0.1 M (for which we previously calculated \(\Delta \psi_\mathrm{m}\) = 40.4 mV and \(\Delta \psi_\mathrm{diff}\) = -0.3 mV). To calculate fluxes we must choose absolute values of the diffusion coefficients, which we take to be \(D_\mathrm{Na}=1.33\cdot 10^{-10}\) m2/s and \(D_\mathrm{Cl}=2.03\cdot 10^{-10}\) m2/s.10
The total NaCl flux is essentially constant throughout the sample (we have adopted a sample length of 10 mm), demonstrating that the imposed linear concentration profile in essence is the steady-state solution. We see that the flux of chloride — which has a much smaller concentration than sodium — is completely dominated by the concentration gradient contribution (eq. 15), while the sodium flux has significant contributions from both the concentration gradient and the electric field (eq. 16). The picture is clear: the faster chloride ions diffuse with a rate set by their individual diffusion coefficient, while the induced electric field boosts the less mobile (but more abundant) sodium ions in order to “keep up”. The behavior is similar to what would be experienced in a bulk solution with a NaCl gradient. In that case, however, the chloride is as much retarded as the sodium is boosted, resulting in a diffusivity for the full salt that is between the two individual values. In compacted bentonite, on the other hand, the electromigration flux is mainly significant for sodium (the counter-ions). Therefore the full salt diffuse with a rate essentially set by the chloride mobility.
We can quantify the relative influcence of electromigration for a specific species by taking the ratio of eqs. 15 and 16, and also utilizing the approximation for \(\nabla \psi (x)\) in eq. 10
This expression explicitly shows that the electromigration flux is negligible for chloride (because \(c^\mathrm{int}_\mathrm{Cl}(x) \ll c_\mathrm{IL}\)), while the \(j_\mathrm{E}/j_\mathrm{conc}\)-ratio for sodium is essentially determined by the factor \(\left (D_\mathrm{Cl}/D_\mathrm{Na} – 1 \right )\) (because \(c^\mathrm{int}_\mathrm{Na}(x) \approx c_\mathrm{IL}\)).
For a similar NaCl-system at lower density, the picture is quite similar (here \(c_\mathrm{IL}\)=1.5 M, with the other parameters as previously; \(\Delta \psi_\mathrm{m}\) = 37.8 mV and \(\Delta \psi_\mathrm{diff}\) = -1.1 mV)
Let’s also play with the relative mobility between anion and cation. In the plot below we have decreased the chloride diffusivity to \(D_\mathrm{Cl}=0.133\cdot 10^{-10}\) m2/s, while keeping with the same values as before for the other parameters (for \(c_\mathrm{IL}\) = 3.0 M).
In this case, the diffusivity factor for the electromigration is \((D_-/D_+ -1) = -0.9\). The negative value indicates an electric field directed in the opposite direction as compared to the previous cases (chloride is now the slower ion). The diffusion potential is \(\Delta \psi_\mathrm{diff}\) = +0.6 mV, which still is quite small, given that the chloride is assumed to have ten times lower mobility as compared with sodium. The evaluated membrane potential is \(\Delta \psi_\mathrm{m}\) = 41.3 mV. We note that the flux is constant throughout the domain and that chloride is completely governed by the concentration gradient contribution, while the sodium flux is composed of two oppositely directed much larger contributions of similar magnitude.
If we instead make the sodium diffusivity 10 times smaller than the chloride diffusivity (\(D_\mathrm{Na}=0.203\cdot 10^{-10}\) m2/s, \(D_\mathrm{Cl}=2.033\cdot 10^{-10}\) m2/s), the effect is more dramatic
With \((D_-/D_+ -1)\) = 9.0, the diffusion potential becomes considerably larger than in any of our previously investigated cases: \(\Delta \psi_\mathrm{diff}\) = -5.2 mV (\(\Delta \psi_\mathrm{m}\) = 35.5 mV). This is reasonable, since a significantly smaller mobility of the counter-ions requires a larger electric field to produce similar fluxes. The sodium flux is therefore dominated by electromigration, and we also note a non-negligible contribution from electromigration for chloride. The overall flux varies quite significantly throughout the clay domain; we’re not really in steady-state.
“Uphill” diffusion
With a solid understanding of the principles for diffusion of a 1:1 electrolyte in compacted bentonite, let’s turn our attention to the “uphill” test (Glaus et al., 2013). The conditions for the main electrolyte (NaClO4) and clay in this study are the same as in our previous case: one reservoir is at concentration 0.1 M, and one at 0.5 M, and the clay is pure high-density Na-montmorillonite. For a further discussion of this study, see this previous post. Here we adopt the value \(c_\mathrm{IL}\) = 4.5 M and set \(c^{(1)}\) = 0.1 M and \(c^{(2)}\) = 0.5 M (in order to have a positive tracer flux). In addition to a main electrolyte, the “uphill” test has a sodium tracer (\(^{22}\mathrm{Na}\)) as a third component, with equal concentration in the two reservoirs (\(10^{-11}\) M). Here we choose sodium diffusivity \(D_\mathrm{Na}=3\cdot 10^{-11}\) m2/s, similar to what is evaluated in Glaus et al. (2007) and Birgersson and Karnland (2009). For perchlorate we choose \(D_\mathrm{ClO_4}=4\cdot 10^{-11}\) m2/s, in order to keep the same diffusivity ratio as for bulk (the perchlorate bulk value is \(1.79\cdot 10^{-9}\) m2/s (Heil, 1995)).
Since the only difference between the “uphill” study and the pure 1:1 case is the presence of a tracer, it is no surprise that concentrations, fluxes and potential behave very similarly
The membrane potential is here \(\Delta \psi_\mathrm{m} = -41.0\) mV, which is essentially identical to what we have evaluated for the NaCl system (the minus sign simply reflects that the left reservoir here has the lower concentration). With a smaller mobility difference between anion and cation, the diffusion potential is even more negligible than for NaCl, \(\Delta \psi_\mathrm{diff} = 0.1\) mV. The tracer flux is not resolved in the above plot, but looks like this
An analytical evaluation of the \(j_\mathrm{E}/J_\mathrm{conc}\)-ratio becomes a bit more complicated for the tracer as compared with the main salt (eq. 17). At the two interfaces, it can be shown that11
We see that this ratio is expected to be small as long as the reservoir concentrations are small as compared with \(c_\mathrm{IL}\). This is also confirmed in the above plot; the electromigration contribution is completely negligible.
This result is the ultimate support for the claim I made in the “uphill” blog post: the phenomena is first and foremost an effect of ordinary Donnan equilibrium, just as conventional cation tracer diffusion is explained. Here we have explicitly included the electric potential in the analysis, and not only showed that electromigration is negligible for the tracer component, but qualitatively described how and where possible effects due to ion mobility differences occur in the set-up as a whole.
Tertre et al. (2024) — the publication critisized in the “uphill” blog post — states that the “uphill” study
[…] demonstrated the marked influence of background electrolyte concentration gradient on tracer diffusion, and thus the necessity to understand the couplings between diffusion of several charged species present at contrasting concentrations and experiencing different concentration gradients.
In contrast to Tertre et al. (2024), we have here presented this understanding and — in contrast to the statement — demonstrated that the “uphill” effect does not depend critically on any additional mechanisms or couplings as compared with conventional cation tracer diffusion.
As for the previous cases, we can play around with the parameters to explore more “extreme” conditions in an “uphill” test. Here we have increased the perchlorate diffusivity to \(D_\mathrm{ClO_4}=9\cdot 10^{-11}\) m2/s (i.e. three times the sodium diffusivity), and lowered \(c_\mathrm{IL}\) to \(c_\mathrm{IL} = \) 2.0 M (corresponding roughly to a density of 1.25 g/cm3).
Predictably, the main counter-ion transport is now dominated by electromigration, and we see some influence of electromigration also on the perchlorate and the sodium tracer. Nevertheless, even for this extreme case, the “uphill” tracer flux is mainly “Fickian” and is principally explaind as an effect due to Donnan equilibrium at the sample interfaces. For this case we have \(\Delta \psi_\mathrm{m}\) = -37.4 mV and \(\Delta \psi_\mathrm{diff}\) = 2.6 mV.
Update (261006):The “uphill” test is modeled with the full Nernst-Planck frameworkhere.
CaCl2
Let’s also examine the pure 2:1 system, which corresponds e.g. to CaCl2 solutions in contact with Ca-montmorillonite. As we discussed in a recent blog post, the Donnan equilibrium for this system is quite different as compared with the 1:1 system. This will naturally also impact the membrane potential. Moreover, experiments indicate that calcium diffuses several times slower than chloride in bentonite (Kozaki et al., 2000), which suggests a larger effect on the diffusion potential. But, similar to previous cases, let’s begin by assuming the same diffusivity ratio as for bulk. With \(D_\mathrm{Cl}=2.03\cdot 10^{-10}\) m2/s, \(D_\mathrm{Ca}=0.79\cdot 10^{-10}\) m2/s, \(c_\mathrm{IL} = \) 3.0 M, and having \(c^{(1)}\) = 0.25 M and \(c^{(2)}\) = 0.05 M12 we get the following picture
The membrane potential is here \(\Delta \psi_\mathrm{m} = 18.8\) mV and the diffusion potential \(\Delta \psi_\mathrm{diff} = -1.1\) mV. Thus, even if the relative influence of the diffusion potential is larger han for NaCl, the main contribution to the membrane potential is still due do Donnan equilibrium; the considerably lower membrane potential in this case (about half of the NaCl value with the same amount of chloride in the reservoirs) is primarily a consequence of Donnan equilibrium: the Donnan factors have in this case a square-root dependence on external concentration. However, as a consequence of calcium mobility being considerably lower than sodium mobility (\(D_-/D_+ – 1\) = 1.57), the magnitude of the diffusion potential is about three times larger than for the NaCl case. The resulting flux is also seen to be not completely constant throughout the clay domain. Nevertheless, the same qualitative picture of the transport mechanism as for NaCl is valid here: chloride transport is dominated by its “Fickian” contribution, diffusing with the rate set by the individual mobility, while the less mobile calcium is boosted by an electromigration contribution, which here is actually larger than its concentration gradient contribution. The salt as a whole diffuses with the rate essentially set by the chloride diffusivity.
Let’s also explore an “extreme” case. Below we have set the Ca mobility to be a factor 10 lower than Cl (\(D_\mathrm{Cl}=2.03\cdot 10^{-10}\) m2/s, \(D_\mathrm{Ca}=0.203\cdot 10^{-10}\) m2/s) and lowered \(c_\mathrm{IL}\) to \(c_\mathrm{IL} = \) 1.5 M.
Here the membrane potential is only \(\psi_\mathrm{m} = 6.7\) mV due to a large contribution from the diffusion potential (\(\Delta \psi_\mathrm{diff} = -12.0\) mV), The calcium flux is seen to be strongly dominated by electromigration, and chloride transport is also strongly affected. We are far from steady-state.
Mixing sodium and calcium
As a final case we consider systems containing both a mono- and a di-valent type of cation. Since cation concentrations are enhanced within the clay, a significant diffusion potential may develop, even for relatively small variations of the external solutions. For instance, with a pure 0.1 M NaCl solution on one side (2), and a mixture of 0.003 M CaCl2 and 0.094 M NaCl on the other (1), we get the following result
Here we have assumed \(c_\mathrm{IL} = \) 3.0 M, \(D_\mathrm{Cl}=2.03\cdot 10^{-10}\) m2/s, \(D_\mathrm{Ca}=0.79\cdot 10^{-10}\) m2/s, and \(D_\mathrm{Na}=1.33\cdot 10^{-10}\) m2/s.
Note that the Donnan equilibrium at the left interface acts as to make the clay contain 50% sodium and 50% calcium (charge wise) even though the calcium content in the reservoir is only 6%! A quite minor “disturbance” in the external solution consequently induce huge transport processes.13 This transport is essentially independent of the presence of an anion, and is governed by mutual diffusion of calcium and sodium. The mobility difference for these two ions thus results in a large diffusion potential: \(\Delta \psi_\mathrm{diff}\) = -5.0 mV. The membrane potential is quite substantial: \(\Delta \psi_\mathrm{m}\) = 11.4 mV. The calcium transport is dominated by the concentration gradient flux, while the oppositely directed sodium flux is more profoundly retarded by the electric field (although it is still dominated by the concentration gradient). We conclude that the full Na-Ca transport process occurs approximately at the rate set by the individual mobility for calcium. This process is essentially charge neutral, i.e. as much Ca charge is transported to the right as Na charge is transported to the left.
It may also be worth noticing that although the chloride concentration is the same in the two external solutions, an internal gradient — and a corresponding chloride flux — is induced
This chloride flux, which is directed to the right and seen to be quite influenced (retarded) by electromigration, is more than two orders of magnitude smaller than the Ca-Na flux. This result implies that also anions may experience seeming “uphill” diffusion (though I’m not sure how easy it would be to detect experimentally).
A lesson here is that calcium concentrations often must be very small in order to be treated as a tracer when added to a sodium system. For example, Tinnacher et al. (2016) incorrectly treats calcium as a tracer when adding 0.001 M CaBr2 to an otherwise pure 0.1 M NaCl-montmorillonite system at 0.8 g/cm3.14 At that density, we may approximate \(c_\mathrm{IL} = 1.0\) M, which gives the following situation (\(D_\mathrm{Cl} = 2.03 \cdot 10^{-10}\) m2/s, \(D_\mathrm{Br}=2.01\cdot 10^{-10}\) m2/s, \(D_\mathrm{Ca}=0.79\cdot 10^{-10}\) m2/s, and \(D_\mathrm{Na}=1.33\cdot 10^{-10}\) m2/s; CaBr2 in reservoir 1)
The amount of calcium in the clay at the left interface is in this case predicted to be about 15% — which certainly cannot be regarded a trace level. Consequently, the main mode of transport in this test is oppositely directed sodium and calcium fluxes that essentially does not involve the anions. In addition, we predict a non-zero membrane potential of \(\Delta \psi_\mathrm{m} = 2.5\) mV, with \(\Delta \psi_\mathrm{diff} = -1.5\) mV.
The bromide, in contrast, can be treated as tracer, and its flux look like this
This flux is about two orders of magnitude smaller than the dominating Ca-Na flux,15 and is seen to be essentially unaffected by the electric field (i.e. mainly “Fickian”).
Additional comments
Here we have investigated transport processes that occur in the presence of chemical gradients between two external reservoirs sandwiching a compacted bentonite component. For this case, compacted bentonite appears to function similar to a conventional charged ion exchange membrane.2 However, compacted bentonite used e.g. for waste storage is quite different from ordinary ion exchange membranes, in that we are more interested in what is going on inside it, rather than directly utilizing its semi-permeable properties. The results discussed in this blog post are useful when interpreting tests that involve bentonite sandwiched between external solutions. For a description of what is going on inside the bentonite component, such results and tests may be very useful, but do not provide a full picture.
As with many topics discussed on the blog, I find the lack of relevant experimental results quite extraordinary. I mean that many labs should routinely measure membrane potentials as part of exploring bentonite systems. Instead, as far as I understand, the study by Yaroschchuck et al. (2007) is the only one published on actual compacted bentonite.16 Moreover, this study appears to be completely forgotten!8
Yaroschchuck et al. (2007) use extremely high density, and it would be interesting with data also at lower densities (1.6 — 1.3 g/cm3, say); the present theory predicts only minor changes in membrane potentials. A “simple” further test of the present description would be to measure the membrane potential in e.g. a pure CaCl2-montmorillonite system. We end this post by presenting such potentials as predicted from the present theory (the calculation uses \(D_\mathrm{Cl} = 2\cdot 10^{-10}\) m2/s and \(D_\mathrm{Ca} = 0.5\cdot 10^{-10}\) m2/s).
Update (260922): The full Nernst-Planck framework is treated within the homogeneous mixture model here.
Footnotes
[1] The
focus here is on what I call
simple ions, i.e. ions that only interact with the bentonite by beeing
part of a diffuse layer. In bentonite, more complex processes are
active for certain species that interact with e.g. montmorillonite
edges. Here
is presented an extension of the homogeneous mixture model for
“truly” sorbing species.
[2]
What is presented in this blog post is, as far as I understand, the
“Teorell-Meyer-Sievers”
model for the membrane potential, applied to compacted
bentonite. This model, which was originally developed in the 1930s,
isusedroutinely within
the ion-exchange membrane scientific discipline. I’m currently reading
up on this literature, and is here using my “usual” terminology
(for bentonite). I have not yet come across sources with specific
focus on bentonite/montmorillonite, and I’m not sure how well
established these are as a membrane materials within this
discipline. In a well-cited review by
Xu (2005),
bentonite is mentioned once — misspelled! Perhaps this is
indicative of a missing link between the ion exchange membrane and
compacted bentonite research fields.
[3] Although different water
chemistries will induce different
swelling pressure repsonses, and thus
couple to mechanical processes in the bentonite, we here ignore
pressure altogether (but we certainly will have reason to return to
this).
[4] Note that
Donnan potentials generally are negative, while
\(\Delta \psi_\mathrm{Donnan}^{(1)}\) and
\(\Delta \psi_\mathrm{Donnan}^{(2)}\) are negative or positive,
depending on whether the contribution decreases or increases the
potential as we move to the right.
[5] Including
activity coefficients, eq. 6 becomes
and the interlayer activity coefficients (\( \gamma^\mathrm{int}\)) will therefore cancel (to the extent that the this quantity can be considered independent of external solution concentration). The contribution from Donnan equilibrium to the membrane potential thus become
[7] The value of \(c_\mathrm{IL}\) (eq. 5) depends on the cation exchange capacity of the material: montmorillonite “from Milos”. Differentsources report this value in the range 0.80 — 0.85 eq./kg.
[8] The obscurity of Yaroshchuck et al. (2007) amazes me. This paper is part of a “trilogy” published in 2007, by the same authors (to a large extent). The other two studies are featured massively on the blog: Glaus et al. (2007) demonstrates that interlayer diffusion dominates mass transfer in compacted bentonite (see here and here), and Van Loon et al. (2007) is the only study that passed our assessment of published chloride equilibrium concentrations in compacted bentonite (see here, here, and here). Glaus et al. (2007) has at the moment 178 citations on Google Scholar, and Van Loon et al. (2007) has 333 citations. Yaroshchuck et al. (2007), in contrast, has 7 citations… Perhaps even more weirdly, Yaroshchuck et al. (2007) is not cited in Van Loon et al. (2007), although the former study presents the framework for Donnan equilibrium between compacted bentonite and a 1:1 electrolyte, and relates it to the findings of Glaus et al. (2007). It should thus had been an “easy win” for Van Loon et al. (2007) to provide a proper explanation for anion exclusion in bentonite, and to make the correct connection between anion and cation tracer through-diffusion. Instead, and for reasons I can’t get my head around, van Loon et al. (2007) get lost in ideas on “anion-accessible porosity”.
[9] Generally, the flux may also have an advective component, which we don’t consider here.
[10] The absolute values of the diffusivities are chosen rather arbitrarily; in this example we have adopted values ten times smaller than the corresponding values in bulk. The absolute value of the corresponding flux is therefore also arbitrary. Here, however, we are mainly interested in how constant the flux is throughout the clay, and in the relative contributions from concentrations and electric fields, rather than the absolute values. In some cases in this section we have adopted empirical values for diffusivities in order to compare also the absolute value of the flux.
[11] The electromigration and concentration gradient fluxes for the tracer in the “uphill” test can be written (see eqs. 15 and 16)
where we have explicitly written the gradients in terms of the internal interface concentrations. Using the relation between Donnan factors and concentrations, this concentration difference ratio can in turn be rewritten
where we have written \(c_\mathrm{tr}\) for the tracer concentration in the reservoirs (it’s equal in the two solutions). The flux ratio can thus be written
[15] The actual measured bromide and calcium fluxes in Tinnacher et al. (2016) differ only by about one order of magnitude. This may be in part due to filter limitations, and in part be an indication that a completely homogeneous treatment begins to break down at these densities.
[16] Some results on bentonite samples of somewhat lower density are presented by Heister (2005).
Here’s an opinion: The compacted bentonite research field is currently
in a terrible state.
After a period away, I’ve recently begun catching up on newly published research in this field. With a fresh perspective, yet still influenced by writing over 30 long-reads over the past years, I can’t help but wonder: what is the problem? Why are a majority of researchers stuck with a view of bentonite1 that essentially makes no sense? And why has this view been the mainstream for decades now?
I get how this might come across: a solitary man ranting on a blog,
criticizing an entire research field in less-than-perfect English. I
probably smell bad and have some wild ideas about why General
Relativity is wrong as well. But what I’m aiming for with this blog
is simply a platform to present an alternative to the mainstream,
primarily because it annoys me as a science-minded person how absurd
this view is.2 I
understand that I will likely struggle to convince anyone who is
already invested in this view, but I’m trying to put myself in the
shoes of e.g. someone entering this field for the first time.
For these reasons, I will try something a little new here: reviewing already published papers. I have touched on this in various forms before, but then usually with a broader topic in mind. Now I intend to critically assess specific publications from the outset. As a first publication to review in this way, I have chosen “Ionic Transport in Nano-Porous Clays with Consideration of Electrostatic Effects” (Tournassat and Steefel, 2015), for the following reasons
It is published in “Reviews in Mineralogy & Geochemistry”, which claims that “The content of each volume consists of fully developed text which can be used for self-study, research, or as a text-book for graduate-level courses.” If anyone aims to learn about ion transport in bentonite from this publication, I would certainly recommend to also consider this review.
It is a quite comprehensive source for many of the claims of the contemporary mainstream view that I have described in earlier blog posts. I guess it makes sense for a publication in “Reviews in Mineralogy & Geochemistry” to reflect the typical view of a research field.
It is published as open access. The article is thus accessible to anyone who wants to check the details.
I will use the abbreviation TS15 in following to refer to this
publication.
Overview
The article covers 38 journal pages (+ references) and includes quite
a lot of topics. At the highest level of headings, the outline look
like this
Introduction (p. 1 — 2)
Classical Fickian Diffusion Theory (p. 2 — 9)
Clay mineral surfaces and related properties (p. 9 — 17)
Constitutive equations for diffusion in bulk, diffuse layer, and interlayer water (p. 17 — 23)
Relative contributions of concentration, activity coefficient and diffusion potential gradients to total flux (p. 24 — 28)
From diffusive flux to diffusive transport equations (p. 28 — 33)
Applications (p. 33 — 37)
Summary and Perspectives (p. 37 — 38)
Given the quite large scope of TS15, I will present this review in
parts, with this first part focusing on the introduction and the
section titled “Classical Fickian Diffusion Theory”.
“Introduction”
I find it remarkable that the authors use terms like “clays” and “clay minerals” when speaking of properties such as “low permeability”, “high adsorption capacity” and “swelling behavior”, and of applications such as nuclear waste storage. I mean that using such general terms here is too broad, as the article focuses solely on systems with swelling/sealing ability. Such an ability is generally connected with a significant cation exchange capacity. Here, I will refer to such systems as “bentonite”, although I am aware that I use the term quite sloppily. But I think this is better than to refer to the components as general “clay minerals” — I don’t think anyone consider it a good idea to e.g. use talc or kaolinite as buffer materials in nuclear waste repositories. Moreover, most of the examples considered in the article are systems that can be described as bentonite. Given the title of the article I also expect a definition of “nano-porous clays”. It is not given here, and the term is actually not used at all in the entire text! (Except one time at the very end.)
After providing a brief overview of the application of (sealing) clay materials, the introduction takes, in my opinion, a rather drastic turn (it happens without even changing paragraphs!).
Clay transport properties are however not simple to model, as they deviate in many cases from predictions made with models developed previously for “conventional” porous media such as permeable aquifers (e.g., sandstone). […] In this respect, a significant advantage of modern reactive transport models is their ability to handle complex geometries and chemistry, heterogeneities and transient conditions (Steefel et al. 2014). Indeed, numerical calculations have become one of the principal means by which the gaps between current process knowledge and defensible predictions in the environmental sciences can be bridged (Miller et al. 2010).
I think the first sentence is too subjective and general. Given the above discussion, here the term “clay transport properties” can cover a million things, if read at face value. Are all of them difficult to model? Also, something does not have to be more difficult just because it deviates from the “convention”. I would argue that several aspects of bentonite actually make it easier to model than, say, sandstone. Advective processes, for example, can often be neglected in compacted bentonite.
I find the statement regarding the advantage of reactive transport
models highly problematic. Not only does it read more like an
advertisement for the authors’ own tools than “fully developed text
for self-study”, but the authors also seem ignorant of issues like
the dangers of overparameterization (a theme that will recur).
“Classical Fickian Diffusion Theory”
As the title of the next section is “Classical Fickian Diffusion Theory,” a reader expects a discussion focused solely on diffusive process, especially when the immediate subtitle reads “Diffusion Basics.” I therefore find it peculiar that this section actually presents the traditional diffusion-sorption model, which describes a combination of diffusion and sorption processes. The model is summarized in eq. 10 in TS15
where \(c\) is the “pore water” concentration of the considered species, \(D_e\) its “effective diffusivity”, \(K_D\) the sorption partition coefficient, \(\rho_d\) dry density, and \(\phi\) porosity.3 For later considerations we also note that TS15 define the denominator on the right hand side as the “rock capacity factor”, \(\alpha = \phi + \rho_dK_D\).
I find it particularly odd that two of the fundamental assumptions of
this specific model are essentially left uncommented, namely that
sorbed ions are immobilized and that the pores contain bulk
water. Instead, the authors appear to question the assumption of
Fickian diffusion in the context of clay systems, i.e. that diffusive
fluxes are assumed proportional to corresponding aqueous concentration
gradients.
This section aims, as far as I can see, to point out shortcomings in the description of diffusion in bentonite, and to motivate further model development. But it should be clear from the outset that using the traditional diffusion-sorption model as the basis for such an endeavor is doomed to fail. The reason for this failure is not due to assuming Fickian diffusion, but due to the other two model assumptions; it has long been demonstrated that exchangeable ions are mobile, and the notion that compacted bentonite contains mainly bulk water is absurd.
After the traditional diffusion-sorption model has been presented, it is evaluated by investigating how it can be fitted to tracer through-diffusion data (this is restatement of original work of Tachi and Yotsjui (2014)). Not surprisingly, it turns out that fitted diffusion coefficients may be unrealistically large. This is of course a direct consequence of the incorrect assumption of immobility in the traditional diffusion-sorption model. TS15 also appear to dismiss the model, saying
This result […] is not physically correct and points out the
inconsistency of the classic Fickian diffusion theory for modeling
diffusion processes in clay media.
I am bothered, though, that they keep using the phrase “classic Fickian diffusion theory”, which inevitably focuses on the Fickian aspect rather than on the obviously incorrect assumptions of the chosen model. Also, rather than simply concluding that the model is incorrect, TS15 continues4
[T]he large changes of \(\mathrm{Cs}^+\) diffusion parameters as a function of chemical conditions (\(D_{e,\mathrm{Cs}^+}\) decreases when the ionic strength increases […]) highlight the need to couple the chemical reactivity of clay materials to their transport properties in order to build reliable and predictive diffusion models.
There is no rationale for such a conclusion. I don’t even completely
understand what “couple the chemical reactivity of clay materials to
their transport properties” mean. Isn’t that what the traditional
diffusion-sorption model attempts? What unrealistic \(D_e\) values
actually highlights is simply that one should not use a model that
assumes immobilization of “sorbed” ions.
However, the experimental observations were completely different: \(^{22}\mathrm{Na}^+\) accumulated in the high NaCl concentration reservoir as it was depleted in the low NaCl concentration reservoir, evidencing non-Fickian diffusion processes.
This is plain wrong. As explained in detail in an earlier post, the diffusion process in the “uphill” test is certainly Fickian. What the test demonstrates is, again, that “sorbed” ions are not immobile.
TS15 also comment on the results of fitting the model to anion tracer through-diffusion data. Here, as is well known, the fitted “rock capacity factor” \(\alpha = \phi + \rho_dK_D\) becomes significantly lower than the porosity \(\phi\). From the perspective of the traditional diffusion-sorption model, this is completely infeasible, as it implies a negative \(K_D\). But rather than simply dismissing the model, TS15 state
The lower \(\alpha\) values for anions than for water indicate that
anions do not have access to all of the porosity.
Also this is incorrect. The porosity5 is an input
parameter rather than a fitting parameter in the traditional
diffusion-sorption model. When claiming that a small value of \(\alpha\)
indicates a decreased porosity, TS15 reinterpret the parameter,
on the fly, in terms of a completely different model:
the effective porosity model. This model has not been mentioned at
all earlier in the article.6
As has been discussed earlier on the blog, the effective porosity model can be fitted to anion tracer through-diffusion data, but now we need to keep track of two different models in the evaluation (something that TS15 do not). Moreover, these two models (the traditional diffusion-sorption and the effective porosity models) are incompatible. But TS15 continue by saying
This result is a first direct evidence of the limitation of the
classic Fickian diffusion theory when applied to clay porous media:
it is not possible to model the diffusion of water and anions with
the same single porosity model. The observation of a lower \(\alpha\)
value for anions than for water led to the development of the
important concept of anion accessible porosity […]
This is a terrible passage. To begin with, the “Fickian” aspect is
also here implied as the problem. But the reason for why the
traditional diffusion-sorption model cannot be fitted to anion tracer
through-diffusion data is of course because this model assumes the
entire pore space to be filled with bulk water. Further, it’s hardly
comprehensible what the authors mean by “it is not possible to model
the diffusion of water and anions with the same single porosity
model”. I think they simply mean that for water you must choose
\(\alpha = \phi\), while for anion through-diffusion you instead must
“choose” \(\alpha < \phi\). But the result \(\alpha < \phi\) should only
lead to the conclusion that the traditional diffusion-sorption model
cannot in any reasonable sense be fitted. A favorable reading of this
passage is to assume that the authors actually mean that the
effective porosity model can only be fitted to anion and water
tracer through-diffusion data by using different values of the
(effective) porosity, and that any “rock capacity factor” should not
appear in this discussion.
Finally, the last sentence gives me headache. Rather than being an “important concept”, I mean that the idea of an “anion accessible porosity” has caused tremendous damage to the development of the bentonite research field for several decades now. We have earlier discussed on the blog that the whole idea of “anion accessible porosity” is based on misunderstandings. We have also demonstrated that the effective porosity model is not valid, even though it can be fitted to anion tracer through-diffusion data. A simple way to see this is to consider closed-cell diffusion data rather than through-diffusion data. Closed-cell tests are simpler than through-diffusion tests, as they don’t involve interfaces between clay and external solutions. We can e.g. take a look at the vast amount of diffusion coefficients for chloride in montmorillonite, presented in Kozaki et al. (1998).
There are in total 55(!) values, corresponding to 55(!) separate tests. These have been systematically varied with respect to density and temperature, but all of them were performed on montmorillonite equilibrated with distilled water. From the perspective of the effective porosity model, the effective porosity in such a system should be minute, perhaps even strictly zero; effective porosities evaluated from chloride through-diffusion tests are well below 1% even at a background concentration as large as 10 mM. Thus, if the idea of “anion accessible porosity” was reasonable, we’d expect extremely low values of the chloride diffusion coefficient in the above plot.7 We’d perhaps also expect a threshold behavior, where chloride diffusivity basically vanishes above a certain density. But this is not at all the behavior: chloride is seen to diffuse just fine in all 55(!) tests, with temperature- and density dependencies that seems reasonable for a homogeneous system. Moreover, chloride behaves very similarly to e.g. sodium, as seen here
Here the sodium data is from Kozaki et al. (1998),8 and it has also been measured in montmorillonite equilibrated with distilled water.
The effective porosity model and the notion of “anion accessible porosity” can consequently be dismissed directly, by comparing with simpler tests than what is done in TS15. The reason that the effective porosity model can be fitted to anion through-diffusion data must be attributed to a misinterpretation of such tests, as they involve also interfaces to external solutions. At least to me it is completely clear that what many researchers interpret as an effective porosity is actually effects of interface equilibrium.
If TS15 were serious about evaluating bentonite diffusion processes in
this section I think they should have done the following:
Discuss the assumptions of ion immobility of sorbed ions and bulk pore water when presenting the traditional diffusion-sorption model. Moreover, they should not call this “Classical Fickian Diffusion Theory”.
Also present and discuss the effective porosity model, as they obviously use it in their evaluations. They actually even seem to promote it! And it is as “Fickian” as the traditional diffusion-sorption model.
Evaluate the models using closed-cell data to avoid misinterpretations arising from complications at bentonite/external solution interfaces.
Conclude that the traditional diffusion-sorption model is not valid for bentonite, and that this is because of the assumptions of immobility of sorbed ions and bulk pore water.
Conclude that the effective porosity model is not valid for bentonite, and that the notion of “anion accessible porosity” is flawed.
Instead, we get a quite confused and incomplete description, mixed
with entirely inaccurate statements. In the end, it is difficult to
understand what the takeaway message of this section really is. A
reader is left with an impression that there is some problem with the
“Fickian” aspect of diffusion, but nothing is spelled out. We have
also been hinted that “anion accessible porosity” is important,
without really having been introduced to the concept/model.
The section ends with the following passage
The limitations of the classic Fickian diffusion theory must find
their origin in the fundamental properties of the clay minerals. In
the next section, these fundamental properties are linked
qualitatively to some of the observations described above.
If “classic Fickian diffusion theory” here is interpreted as “the traditional diffusion-sorption model” (which is literally what has been presented), the first sentence is both incorrect and trivial at the same time. The traditional diffusion-sorption model does not have “limitations” — it is fundamentally incorrect as a model for bentonite. The reason for this is that exchangeable ions are not immobile and that bentonite does not contain significant amounts of bulk water. Both of these reasons can be linked to “fundamental properties” of some specific clay minerals.
But it is clear that TS15 also have vaguely promoted the concept of “anion accessible porosity” and the effective porosity model. Are these not included in “the classic Fickian diffusion theory”? If not, why then is a model that assumes sorption and immobilization?
How can it not be immediately obvious to everyone that the
diffusion process is much simpler than the contemporary descriptions?
As we have brought up the data from Kozaki etal. (1998), I would like to end this blog post by further considering actual profiles of chloride and sodium diffusing in montmorillonite.
This figure shows the corresponding normalized concentration profiles after 23.7 hours in closed-cell tests performed at \(50\;^\circ\mathrm{C}\) in Na-montmorillonite at dry density \(1.8 \;\mathrm{g/cm^3}\) that has been equilibrated with distilled water. In the case of sodium, both the profile evaluated from Fick’s second law (orange line) and measured values (circles) are plotted. In the case of chloride, no measured values are available, but the value of the diffusion coefficient is the result of fitting Fick’s second law (green line) to such data.
From the perspective of the traditional diffusion-sorption model, the
sodium profile is supposed to represent the combined result of ions
diffusing in bulk water, at a rate many orders of magnitude larger
than in pure water, while being strongly retarded due to sorption onto
“the solid” (where the ions are immobile). This is clearly nonsense,
and something that I think TS15 actually tries to communicate.
From the perspective of the effective porosity model, on the other
hand, the chloride profile is supposed to be the result of the ion
diffusing in an essentially infinitesimal fraction of the pore volume,
which magically is perfectly interconnected in all samples on which such
tests are conducted. This is of course just as nonsensical as
the above interpretation of the sodium profile, but in this case TS15
appear to promote the model (the “important concept of anion
accessible porosity”).
Note that these two simple ions, at the end of the day, diffuse very similarly (please stop reading for a moment and contemplate the above plot). If sodium and chloride actually migrate in completely different domains and are subject to completely different physico-chemical processes, this “coincident” would be more than a little weird. Especially given that the two ions show similar diffusive behavior across a wide range of densities. To me, this simple observation makes it evident that ion diffusion in bentonite at the basic level is much simpler than what is suggested by the contemporary mainstream view. I mean that it is completely obvious that all ions in bentonite diffuse in the same type of quite homogeneous domain. And since it cannot be argued that the pore volume is dominated by anything other than interlayers at 1.8 g/cm³, this homogeneous domain is the interlayer domain at any relevant density. The evidence has been available for at least 25 years (in fact much longer than that). How can this be difficult to grasp?
Update (250213): Part II of this review is found here.
Footnotes
[1] By “bentonite” I here mean any type of smectite-rich system with a significant cation exchange capacity.
[2] The irony is that the “alternative” in a
broader perspective is more mainstream than the “mainstream”
view. I basically propose to obey the laws of thermodynamics.
[3] I have simplified the notation here somewhat compared with how it is written in TS15. As many others, TS15 call this equation “Fick’s second law” (via their eq. 4), which is not correct. Fick’s laws refer strictly to pure diffusion processes. However, the equation has the same form as Fick’s second law, if \(D_e/(\phi + \rho_d K_D)\) is treated as a single constant (often referred to as the apparent diffusivity).
[4] This behavior is of course not
unique for cesium; I don’t know why TS15 focus so hard on that ion
here.
[5] “Porosity” is a volume ratio. I’m not a fan of that the word has also begun to mean “pore space” in the bentonite scientific literature.
[6]
In fact, \(\alpha\) has earlier in the article been unambiguously
related to sorption:
If the species \(i\) is also adsorbed on or
incorporated into the solid phase, then it is possible to define a
rock capacity factor \(\alpha_i\) that relates the concentration in
the porous media to the concentration in solution
[7] That the diffusivity is much too large for an effective porosity interpretation to make sense can also be seen from invoking Archie’s law, which is quite popular in bentonite scientific papers.
Here \(D_0\) is the diffusivity in pure bulk water, which is about \(2\cdot 10^{-9} \;\mathrm{m^2/s}\) for chloride. Using the popular choice \(n \approx 2\) and choosing e.g. \(\epsilon_\mathrm{eff} = 0.001\) (most probably an overestimation when using distilled water), we get
This is more than twenty times the actual value for \(D_0\). (\(D_e = 5.1\cdot 10^{-14} \;\mathrm{m^2/s}\) is evaluated from Kozaki’s data at \(1.4 \;\mathrm{g/cm^3}\) and \(25\;^\circ\mathrm{C}\))
[8] Note! This publication is different from the
chloride study.
The paper follows a structure recognizable from several others that we have considered previously on the blog: It starts off with an introduction section containing several incorrect or unfounded statements1 regarding bentonite.2 It then presents some experimental results that makes it evident that no real progress has been made for a long time regarding e.g. experimental design.3 The major part of the paper is devoted to a “results and discussion” section with several incorrect statements and inferences, speculation, and irrelevant modeling.
[I]nfluence of a background electrolyte concentration gradient on the diffusion of anionic and cationic species at trace concentrations has […] been rarely investigated. Notable exceptions are the DR-A in situ diffusion experiment conducted at the Mont-Terri laboratory (Soler et al., 2019), and an “uphill” diffusion experiment of a \(^{22}\mathrm{Na}^+\) tracer in a compacted sodium montmorillonite (Glaus et al., 2013). These two studies demonstrated the marked influence of background electrolyte concentration gradient on tracer diffusion, and thus the necessity to understand the couplings between diffusion of several charged species present at contrasting concentrations and experiencing different concentration gradients. The experiment from Glaus et al. (2013) also demonstrated the importance of considering diffusion processes occurring in the porosity next to the charged surface of clay minerals (i.e., the porosity associated to the EDL of particles).
This quotation contains two statements relating to Glaus et al. (2013), both of which I think are problematic4
It basically claims that the “uphill” phenomenon is due to diffusive couplings between several types of ions. Of course, ion diffusion always involves couplings between different types of ions, due to the requirement of electroneutrality. But it is clear that Tertre et al. (2024) mean that the “uphill” effect is caused by additional couplings that are not present in chemically homogeneous systems.
It says that Glaus et al. (2013) demonstrates the importance to consider diffuse layers. I agree with this, but it is written in a way that implies that there also are other relevant “porosities”, and that there are other types of tests where ion diffusion in bentonite is not significantly influenced by the presence of diffuse layers.
As one of the authors of the “uphill” study, I would here like to argue for why I think the above statements are problematic and give some background context.
The “uphill” diffusion experiment
The “uphill” study actually originated from a prediction presented by me in a conference poster session. This poster discussed the role of the quantity \(D_e\), using the exact same theory that we had previously used to explain the diffusive behavior of tracer ions in compacted bentonite as an effect of Donnan equilibrium in a homogeneous system. In particular, it pointed out that \(D_e\) — although universally referred to as the (effective) “diffusion coefficient” — is not a diffusion coefficient in the context of compacted bentonite. I have continued this discussion in laterpapers, and in several postson this blog.
In the poster, we suggested the “uphill” experiment as a demonstration of the shortcoming of \(D_e\). If the two reservoirs in a through-diffusion test are maintained at different background concentrations, the theory predicts a non-zero tracer flux for a vanishing external tracer concentration difference, i.e. an “infinite” value of \(D_e\). The suggestion caught the interest of an experimental group, and after a successful collaboration we could present the results of an actual “uphill” experiment. Without making too much of an exaggeration, I would say that the results of this experiment were basically exactly as predicted.
Given this background, it should be clear that the tests in Glaus et al. (2013) follow exactly the same rules as tests in chemically homogeneous systems, rather than demonstrating “the necessity to understand the couplings between diffusion of several charged species present at contrasting concentrations”. Although it is quite clearly stated already in the abstract in Glaus et al. (2013), there is apparently still a need to communicate this explanation. Let me therefore try that here.
The “uphill” diffusion phenomenon explained
Consider an ordinary aqueous solution containing radioactive \(^{22}\mathrm{Na}\) and stable \(^{23}\mathrm{Na}\). The fraction of \(^{22}\mathrm{Na}\) ions can be written \(c_\mathrm{ext}/C_\mathrm{bkg}\), where \(c_\mathrm{ext}\) is the \(^{22}\mathrm{Na}\) concentration, and \(C_\mathrm{bkg}\) is the total sodium concentration (the “tracer” and “background” concentrations, respectively).
Since \(^{23}\mathrm{Na}\) and \(^{22}\mathrm{Na}\) are basically chemically indistinguishable, the same \(^{22}\mathrm{Na}\)-fraction will be maintained in any system with which this solution is in equilibrium. In particular, if the solution is in equilibrium with a montmorillonite interlayer solution, we can write
where \(c_\mathrm{int}\) and \(C_\mathrm{int}\) are the \(^{22}\mathrm{Na}\) and total interlayer concentrations, respectively. The total interlayer cation concentration (\(C_\mathrm{int}\)) can be handled in different ways, but it is important to note that this is a substantial number under all conditions, relating to the cation exchange capacity.5 Rearranging eq. 1 gives
Since the interlayer cation concentration is always larger than the corresponding background concentration, the above equation tells us that the corresponding interlayer tracer concentration becomes enhanced, by the factor \(C_\mathrm{int}/C_\mathrm{bkg}\).
Conventional through-diffusion
This enhancement mechanism causes the diffusional behavior of \(^{22}\mathrm{Na}\) in conventional through-diffusion experiments in bentonite. In such experiments, the tracer concentration in the target reservoir is usually kept near zero, and the actual steady-state concentration gradient in the interlayers is
where we have indexed the tracer concentration in the source reservoir with “\((1)\)”, labeled the sample length \(L\), and assumed that ions diffuse in the \(x\)-direction. The corresponding flux is thus (Fick’s law)
where \(D_c\) denotes the (macroscopic) diffusivity in the interlayers, and \(\phi\) is porosity. Keeping \(c_\mathrm{ext}^{(1)}\) constant, eq. 2 shows that the \(^{22}\mathrm{Na}\) steady-state flux increases indefinitely as the background concentration is made small, in full agreement with experimental observation.6
The picture below illustrates the concentration conditions in an conventional through-diffusion test.
Here we have chosen \(C_\mathrm{int}=\) 4.0 M, the background concentration in the two reservoirs (blue) is put equal to 0.1 M, and the tracer concentration (orange) is put to 0.1 mM in reservoir 1 (and zero i reservoir 2). The corresponding internal tracer gradient is plotted in the right side diagram, and the resulting diffusive flux is indicated by the arrow.
“Uphill” diffusion
To explain the “uphill” effect the only modifications needed in the above derivation is to allow for different background concentrations in the external reservoirs, and to recognize that the tracer concentration in the clay on the “target” side (indexed “\((2)\)”) no longer is zero. Considering the tracer concentration enhancement at both interfaces, the steady-state interlayer concentration gradient then reads
To be more concrete, let’s assume that \(C_\mathrm{bkg}^{(2)} = 5\cdot C_\mathrm{bkg}^{(1)}\), which is the same ratio as in Glaus et al. (2013). We then have
Note that we recover the conventional through-diffusion result (eq. 2) from this expression, if we put \(c_\mathrm{ext}^{(2)}= 0\). But if we e.g. set the tracer concentration equal in both reservoirs, we still have a flux from side \((1)\) to side \((2)\), of size \(j = 4/5 \cdot \phi D_c\cdot C_\mathrm{int}/C_\mathrm{bkg}^{(1)}\cdot c_\mathrm{ext}^{(1)}\). And even if we make \(c_\mathrm{ext}^{(2)}\) larger than \(c_\mathrm{ext}^{(1)}\) — as long as \(c_\mathrm{ext}^{(1)}< c_\mathrm{ext}^{(2)} < 5\cdot c_\mathrm{ext}^{(1)}\) — we still have a diffusive flux from side \((1)\) to side \((2)\), i.e seeming “uphill” diffusion.
Below is illustrated the concentration conditions in an “uphill” configuration.
In contrast to the above illustration for conventional through-diffusion, the background concentration in reservoir 2 is here raised to 0.5 M and the tracer concentration in reservoir 2 is put equal to 0.2 mM. We see that, although tracers are transported to the reservoir with higher concentration, the process is still ordinary Fickian diffusion, as the internal tracer gradient has the same direction as in the conventional case.
We can now conclude what was stated above: The “uphill” diffusion effect is caused by exactly the same mechanism that cause the behavior of cation diffusion in conventional bentonite through-diffusion tests. This mechanism is ion equilibrium between clay and external solutions at the two interfaces. In this particular case, with sodium tracers diffusing in a sodium background, we don’t need to invoke the full ion equilibrium framework in order to quantify the fluxes, but can rely on the very robust result that any two systems in equilibrium have the same tracer fraction (eq. 1).
Reexamining the Tertre et al. (2024) statements
With the explanation for the “uphill” effect established, let’s re-examine the problematic statements in Tertre et al. (2024) identified above
Glaus et al. (2013) cannot be used to support a claim of “marked influence” of additional diffusional couplings. The opposite is true: Glaus et al. (2013) found no significant influence from mechanisms beyond those in chemically homogeneous conditions.
The “uphill” effect was predicted from taking the idea seriously that diffusion in compacted bentonite is fully governed by interlayer properties. Singling out Glaus et al. (2013) as the study that demonstrates the importance of diffuse layers7 therefore gives the wrong impression. Rather, what Glaus et al. (2013) demonstrates, in conjunction with corresponding conventional through-diffusion results, is that compacted bentonite contains insignificant amounts of bulk water (what Tertre et al. (2024) call “interparticle water”).
A way forward (if anybody cares)
After the uphill study was published I was for a while under the illusion that things would begin to change within the compacted bentonite research field. Not only did the study, to my mind, deal a fatal blow to any bentonite model that relies on the presence of a bulk water phase in the clay. It also opened up a whole new area of interesting studies to conduct. Now, some 11 years later, I can disappointingly conclude that not a single additional study has been presented that explore the ideas here discussed.8 And, regarding bentonite models, bulk water is apparently alive and kicking, as has been discussed ad nauseum on this blog.
Experimentally, there are a number of interesting questions looking for answers. In particular, we actually do expect additional mechanisms to play a role in chemically inhomogeneous systems, e.g. osmosis, and other effects due to presence of salt concentration gradients and electrostatic potential differences. It may be argued for why such effects are not significant in Glaus et al. (2013), but it is of course both of fundamental and practical interest to understand under which conditions they are. The original “uphill” study is e.g. performed at quite extreme density (\(1900\;\mathrm{m^3/kg}\)). How would the result differ at \(1600\;\mathrm{m^3/kg}\) or \(1300\;\mathrm{m^3/kg}\)? Also, how would the results change with other choices of the reservoir concentrations, and how would the results differ if one of the cations is not at trace level (e.g. a system with comparable amounts of sodium and potassium)?
Even under the conditions of the original study, there are several predictions left to verify. If e.g. \(c^{(1)}_\mathrm{ext} = c^{(2)}_\mathrm{ext}/5\), the theory predicts zero flux (implying \(D_e = 0\)). The theory also implies that when performing “conventional” through-diffusion, the actual level of the background concentration in the target reservoir is irrelevant, as long as the tracer concentration is kept at zero.
In fact, one can imagine making a whole cycle of through-diffusion tests to explore the ideas here discussed, as illustrated in this animation
The resulting steady-state flux for various external conditions is indicated by the arrow. Here, the full ion equilibrium framework was used to calculate the internal concentrations (giving an internal gradient also in \(C_\mathrm{int}\)). Background concentrations and total interlayer concentration is chosen to be comparable with Glaus et al. (2013), while the choice for tracer concentration is arbitrary.9
With the risk of sounding hubristic, the number of experiments suggested in the above animation could have given enough material for several Ph.D. theses. But here we are, in the year 2024, without even a replication of the “uphill” effect. Instead, a basically entire research field has been stuck for decades with the ludicrous idea that models of compacted bentonite should be based on a bulk water description. I find this both hilarious and horrific.
Update (260803):A further treatment of the “uphill” test with focus on the electrostatic potential is found here. Update (261006):The “uphill” test is modeled with the full Nernst-Planck framework and the homogeneous mixture modelhere.
Footnotes
[1] For example (follow links to discussions on these issues):
Related to using the traditional diffusion-sorption model, it assumes \(D_e\) to be a real diffusion coefficient, which it is not. I find this particularly remarkable in a paper that deals with the presence of “saline gradients”. A motivation behind e.g. the “uphill” test is to point out the shortcomings of \(D_e\), as discussed in the rest of this blog post.
It claims that “anionic and cationic tracers do not experience the same overall accessible porosity”, which is unjustified.
It claims that “diffusion rates” of anions are decreased and “diffusion rates” of cations are increased, compared to “neutral species”, due to different interactions with diffuse layers. But this is not true generally.
[2] I use the word “bentonite” here quite loosely. Tertre et al. (2024) use wordings such as “clayey samples”, “argillaceous rocks” and “clayey formation”, but it is clear that the presented material is supposed to apply to actual bentonite.
[3] I’m specifically thinking about that cation tracer through-diffusion tests at low background concentration is not a good idea, and that it is completely clear from the results presented in Tertre et al. (2024) that some of these are mainly controlled by diffusion in the confining filters. Estimating a “rock capacity factor” larger than 750 for sodium tracers in a sodium-clay (at 20 mM background concentration) should have set off all alarm bells.
[4] Regarding Soler et al. (2019), I think that whole study is problematic, which I might argue for in a separate blog post.
[5] Glaus et al. (2013) invoke the “exchange site” activity \([\mathrm{NaX}]\) to discuss this quantity. I personally prefer relating it to the quantity \(c_\mathrm{IL}\) that is defined within the homogeneous mixture model.
[6] This agreement has been shown to be quantitative, see e.g. Glaus et al. (2007), Birgersson and Karnland (2009) and Birgersson (2017). Note that this result is quite independent of how many “porosities” you choose to include in a model; it’s merely a consequence of treating the dominating pores (interlayers) adequately. Further, note that measuring the diverging fluxes in the limit of low background concentration becomes increasingly difficult, as the confining filters becomes rate limiting.
[7] In the present context, I presume the terms “diffuse layer” and “interlayer” to be more or less equivalent. Other authors instead make an unjustified distinction, that I have addressed here.
[8] There are afewexamples of published studies where effects of the kind discussed here are present, but where the authors don’t seem to be aware of it.
[9] Tracer concentrations in Glaus et al. (2013) is much smaller, but this value does not affect any behavior, as long as it is small in comparison with total concentration.
What do authors mean when they say that bentonite has semi-permeable properties? Take for example this statement, from Bradbury and Baeyens (2003)1
[…] highly compacted bentonite can function as an efficient semi-permeable membrane (Horseman et al., 1996). This implies that the re-saturation of compacted bentonite involves predominantly the movement of water molecules and not solute molecules.
Judging from the reference to Horseman et al. (1996) — which we look at below — it is relatively clear that Bradbury and Baeyens (2003) allude to the concept of salt exclusion when speaking of “semi-permeability” (although writing “solute molecules”). But a lowered equilibrium salt concentration does not automatically mean that salt is less transferable.
A crucial question is what the salt is supposed to permeate. Note that a semi-permeable component is required for defining both swelling pressure and salt exclusion. In case of bentonite, this component is impermeable to the clay particles, while it is fully permeable to ions and water (in a lab setting, it is typically a metal filter). But Bradbury and Baeyens (2003) seem to mean that in the process of transferring aqueous species between an external reservoir and bentonite, salt is somehow effectively hindered to be transferred. This does not make much sense.
Consider e.g. the process mentioned in the quotation, i.e. to
saturate a bentonite sample with a salt solution. With
unsaturated bentonite, most bets are off regarding Donnan equilibrium,
and how salt is transferred depends on the details of the saturation
procedure; we only know that the external and internal salt
concentrations should comply with the rules for salt exclusion once
the process is finalized.
Imagine, for instance, an unsaturated sample containing bentonite
pellets on the cm-scale that very quickly is flushed with the
saturating solution, as illustrated in this state-of-the-art,
cutting-edge animation
The evolution of the salt concentration in the sample will look
something like this
Initially, as the saturating solution flushes the sample, the
concentration will be similar to that of the external concentration
(\(c_\mathrm{ext}\)). As the sample reaches saturation, it contains more
salt than what is dictated by Donnan equilibrium (\(c_\mathrm{eq.}\)),
and salt will diffuse out.
In a process like this it should be obvious that the bentonite not in any way is effectively impermeable to the salt. Note also that, although this example is somewhat extreme, the equilibrium salt concentration is probably reached “from above” in most processes where the clay is saturated with a saline solution: too much salt initially enters the sample (when a “microstructure” actually exists) and is later expelled.
Also for mass transfer between an external solution and an already saturated sample does it not make sense to speak of “semi-permeability” in the way here discussed. Consider e.g. a bentonite sample initially in equilibrium with an external 0.3 M NaCl solution, where the solution suddenly is switched to 1.0 M. Salt will then start to diffuse into the sample until a new (Donnan) equilibrium state is reached. Simultaneously (a minute amount of) water is transported out of the clay, in order for the sample to adapt to the new equilibrium pressure.2
There is nothing very “semi-permeabilic” going on here — NaCl is
obviously free to pass into the clay. That the equilibrium clay
concentration in the final state happens to be lower than in the
external concentration is irrelevant for how how difficult it is to
transfer the salt.
But it seems that many authors somehow equate “semi-permeability” with salt exclusion, and also mean that this “semi-permeability” is caused by reduced mobility for ions within the clay. E.g. Horseman et al. (1996) write (in a section entitled “Clays as semi-permeable membranes”)
[…] the net negative electrical potential between closely spaced clay particles repel anions attempting to migrate through the narrow aqueous films of a compact clay, a phenomenon known as negative adsorption or Donnan exclusion. In order to maintain electrical neutrality in the external solution, cations will tend to remain with their counter-ions and their movement through the clay will also be restricted (Fritz, 1986). The overall effect is that charged chemical species do not move readily through a compact clay and neutral water molecules may be able to pass more freely.
It must be remembered that Donnan exclusion occurs in many systemsother than “compact clay”. By instead considering e.g. a ferrocyanide solution, it becomes clear that salt exclusion has nothing to do with how hindered the ions are to move in the system (as long as they move). KCl is, of course, not excluded from a potassium ferrocyanide system because ferrocyanide repels chloride, nor do such interactions imply restricted mobility (repulsion occurs in all salt solutions). Similarly, salt is not excluded from bentonite because of repulsion between anions and surfaces (also, a negative potential does not repel anything — charge does).
In the above quotation it is easy to spot the flaw in the argument by switching roles of anions and cations; you may equally incorrectly say that cations are attracted, and that anions tag along in order to maintain charge neutrality.
The idea that “semi-permeability” (and “anion” exclusion) is
caused by mobility restrictions for the ions within the
bentonite, while water can “pass more freely” is found in many
places in the bentonite
literature. E.g. Shackelford and Moore (2013) write (where, again, potentials are
described as repelling)
In [the case of bentonite], when the clay is compressed to a sufficiently high density such that the pore spaces between adjacent clay particles are minimized to the extent that the electrostatic (diffuse double) layers surrounding the particles overlap, the overlapping negative potentials repel invading anions such that the pore becomes excluded to the anion. Cations also may be excluded to the extent that electrical neutrality in solution is required (e.g., Robinson and Stokes, 1959).
This phenomenon of anion exclusion also is responsible for the existence of semipermeable membrane behavior, which refers to the ability of a porous medium to restrict the migration of solutes, while allowing passage of the solvent (e.g., Shackelford, 2012).
[…] TOT layers bear a negative structural charge that is compensated by cation accumulation and anion depletion near their surfaces in a region known as the electrical double layer (EDL). This property gives clay materials their semipermeable membrane properties: ion transport in the clay material is hindered by electrostatic repulsion of anions from the EDL porosity, while water is freely admitted to the membrane.
and Tournassat and Steefel (2019) write (where, again, we can switch roles of “co-” and “counter-ions”, to spot one of the flaws)
The presence of overlapping diffuse layers in charged nanoporous media is responsible for a partial or total repulsion of co-ions from the porosity. In the presence of a gradient of bulk electrolyte concentration, co-ion migration through the pores is hindered, as well as the migration of their counter-ion counterparts because of the electro-neutrality constraint. This explains the salt-exclusionary properties of these materials. These properties confer these media with a semi-permeable membrane behavior: neutral aqueous species and water are freely admitted through the membrane while ions are not, giving rise to coupled transport processes.
I am quite puzzled by these statements being so commonplace.3 It does not surprise me that all the quotations basically state some version of the incorrect notion that salt exclusion is caused by electrostatic repulsion between anions and surfaces — this is, for some reason, an established “explanation” within the clay literature.4 But all quotations also state (more or less explicitly) that ions (or even “solutes”) are restricted, while water can move freely in the clay. Given that one of the main features of compacted bentonite components is to restrict water transport, with hydraulic conductivities often below 10-13 m/s, I don’t really know what to say.
Furthermore, one of the most investigated areas in bentonite research is the (relatively) high cation transport capacity that can be achieved under the right conditions. In this light, I find it peculiar to claim that bentonite generally impedes ion transport in relation to water transport.
Bentonite as a non-ideal semi-permeable membrane
As far as I see, authors seem to confuse transport between external
solutions and clay with processes that occur between two
external solutions separated by a bentonite component. Here is
an example of the latter set-up
The difference in concentration between the two solutions implies
water transport — i.e. osmosis — from the reservoir with lower salt
concentration to the reservoir with higher concentration. In this
process, the bentonite component as a whole functions as the membrane.
The bentonite component has this function because in this process it
is more permeable to water than to salt (which has a driving force to
be transported from the high concentration to the low concentration
reservoir). This is the sense in which bentonite can be said to be
semi-permeable with respect to water/salt. Note:
Salt is still transported through the bentonite. Thus, the bentonite component functions fundamentally only as a non-ideal membrane.
Zooming in on the bentonite component in the above set-up, we note that the non-ideal semi-permeable functionality emerges from the presence of two ideal semi-permeable components. As discussed above, the ideal semi-permeable components (metal filters) keep the clay particles in place.
The non-ideal semi-permeability is a consequence of salt exclusion. But these are certainly not the same thing! Rather, the implication is: Ideal semi-permeable components (impermeable to clay) \(\rightarrow\) Donnan effect \(\rightarrow\) Non-ideal semi-permeable membrane functionality (for salt)
The non-ideal functionality means that it is only relevant during non-equilibrium. E.g., a possible (osmotic) pressure increase in the right compartment in the illustration above will only last until the salt has had time to even out in the two reservoirs; left to itself, the above system will eventually end up with identical conditions in the two reservoirs. This is in contrast to the effect of an ideal membrane, where it makes sense to speak of an equilibrium osmotic pressure.
None of the above points depend critically on the membrane material being bentonite. The same principal functionality is achieved with any type of Donnan system. One could thus imagine replacing the bentonite and the metal filters with e.g. a ferrocyanide solution and appropriate ideal semi-permeable membranes. I don’t know if this particular system ever has been realized, but e.g. membranes based on polyamide rather than bentonite seems more commonplace in filtration applications (we have now opened the door to the gigantic fields of membrane and filtration technology). From this consideration it follows that “semi-permeability” cannot be attributed to anything bentonite specific (such as “overlapping double layers”, or direct interaction with charged surfaces).
I think it is important to remember that, even if bentonite is semi-permeable in the sense discussed, the transfer of any substance across a compacted bentonite sample is significantly reduced (which is why we are interested in using it e.g. for confining waste). This is true for both water and solutes (perhaps with the exception of some cations under certain conditions).
“Semi-permeability” in experiments
Even if bentonite is not semi-permeable in the sense described in many
places in the literature, its actual non-ideal semi-preamble
functionality must often be considered in compacted clay
research. Let’s have look at some relevant cases where a bentonite
sample is separated by two external solution reservoirs.
The traditional tracer through-diffusion test maintains identical
conditions in the two reservoirs (the same chemical compositions and
pressures) while adding a trace amount of the diffusing substance to
the source reservoir. The induced tracer flux is monitored by
measuring the amount of tracer entering the target reservoir.
In this case the chemical potential is identical in the two reservoirs for all components other than the tracer, and no additional transport processes are induced. Yet, it should be kept in mind that both the pressure and the electrostatic potential is different in the bentonite as compared with the reservoirs. The difference in electrostatic potential is the fundamental reason for the distinctly different diffusional behavior of cations and anions observed in these types of tests: as the background concentration is lowered, cation fluxes increase indefinitely (for constant external tracer concentration) while anion fluxes virtually vanish.
Tracer through-diffusion is often quantified using the parameter
\(D_e\), defined as the ratio between steady-state flux and
the external concentration
gradient.5 \(D_e\) is thus a
type of ion permeability coefficient, rather than a diffusion
coefficient, which it nevertheless
often is assumed to be.
Typically we have that
\(D_e^\mathrm{cation} > D_e^\mathrm{water} > D_e^\mathrm{anion}\) (where
\(D_e^\mathrm{cation}\) in principle may become
arbitrary large). This behavior both demonstrates the underlying
coupling to electrostatics, and that “charged chemical species”
under these conditions hardly can be said to move less readily through
the clay as compared with water molecules.
Measuring hydraulic conductivity
A second type of experiment where only a single component is
transported across the clay is when the reservoirs contain pure water
at different pressures. This is the typical set-up for measuring the
so-called hydraulic conductivity of a clay
component.6
Even if no other transport processes are induced (there is nothing
else present to be transported), the situation is here more complex
than for the traditional tracer through-diffusion test. The difference
in water chemical potential between the two reservoirs implies a
mechanical coupling to the clay, and a
corresponding response in density distribution. An inhomogeneous
density, in turn, implies the presence of an electric field. Water
flow through bentonite is thus fundamentally coupled to both
mechanical and electrical processes.
In analogy with \(D_e\), hydraulic conductivity is defined as the ratio
between steady-state flow and the external pressure
gradient. Consequently, hydraulic conductivity is an effective mass
transfer coefficient that don’t directly relate to the fundamental
processes in the clay.
An indication that water flow through bentonite is more subtle than what it may seem is the mere observation that the hydraulic conductivity of e.g. pure Na-montmorillonite at a porosity of 0.41 is only 8·10-15 m/s. This system thus contains more than 40% water volume-wise, but has a conductivity below that of unfractioned metamorphic and igneous rocks! At the same time, increasing the porosity by a factor 1.75 (to 0.72), the hydraulic conductivity increases by a factor of 75! (to 6·10-13 m/s7)
Mass transfer in a salt gradient
Let’s now consider the more general case with different chemical
compositions in the two reservoirs, as well as a possible pressure
difference (to begin with, we assume equal pressures).
Even with identical hydrostatic pressures in the reservoirs, this configuration will induce a pressure response, and consequently a density redistribution, in the bentonite. There will moreover be both an osmotic water flow from the right to the left reservoir, as well as a diffusive solute flux in the opposite direction. This general configuration thus necessarily couples hydraulic, mechanical, electrical, and chemical processes. Update (260803):The electrostatic potential is treated here.
This type of configuration is considered e.g. in the study of osmotic effects in geological settings, where a clay or shale formation may act as a membrane.8 But although this configuration is highly relevant for engineered clay barrier systems, I cannot think of very many studies focused on these couplings (perhaps I should look better).
For example, most through-diffusion studies are of the tracer type discussed above, although evaluated parameters are often used in models with more general configurations (e.g. with salt or pressure gradients). Also, I am not aware of any measurements of hydraulic conductivity in case of a salt gradient (but the same hydrostatic pressure), and I am even less aware of such values being compared with those evaluated in conventional tests (discussed previously).
A quite spectacular demonstration that mass transfer may occur very differently in this general configuration is the seeming steady-stateuphill diffusion effect: adding an equal concentration of a cation tracer to the reservoirs in a set-up with a maintained difference in background concentration, a tracer concentration difference spontaneously develops. \(D_e\) for the tracer can thus equal infinity,9 or be negative (definitely proving that this parameter is not a diffusion coefficient). I leave it as an exercise to the reader to work out how “semi-permeable” the clay is in this case. Update (240822):The “uphill” diffusion effect is further discussed here.
A process of practical importance for engineered clay barrier systems
is hyperfiltration of salts. This process will occur when a sufficient
pressure difference is applied over a bentonite sample contacted with
saline solutions. Water and salt will then be transferred in the same
direction, but, due to exclusion, salt will accumulate on the
inlet side. A steady-state concentration profile for such a process
may look like this
The local salt concentration at the sample interface on the inlet side
may thus be larger than the concentration of the injected
solution. This may have consequences e.g. when evaluating hydraulic
conductivity using saline solutions.
Hyperfiltration may also influence the way a sample becomes saturated, if saturated with a saline solution. If the region near the inlet is virtually saturated, while regions farther into the sample still are unsaturated, hyperfiltration could occur. In such a scenario the clay could in a sense be said to be semi-permeable (letting through water and filtrating salts), but note that the net effect is to transfer more salt into the sample than what is dictated by Donnan equilibrium with the injected solution (which has concentration \(c_1\), if we stick with the figure above). Salt will then have to diffuse out again, in later stages of the process, before full equilibrium is reached. This is in similarity with the saturation process that we considered earlier.
[2] This is more than a thought-experiment; a test just like this was conducted by Karnland et al. (2005). Here is the recorded pressure response of a Na-montmorillonite sample (dry density 1.4 g/cm3) as it is contacted with NaCl solutions of increasing concentration
[3] As a side note, is the region near the surface supposed to be called “diffuse layer”, “electrical double layer”, or “electrostatic (diffuse double) layer”?
[5] This is not a gradient in the mathematical sense, but is defined as \( \left (c_\mathrm{target} – c_\mathrm{source} \right)/L\), where \(L\) is sample length.
[6] Hydraulic conductivity is often also measured
using a saline solution, which is commented on below.
[7] Which
still is an a amazingly small hydraulic conductivity, considering
the the water content.
[9] Mathematically, the statement “equal infinity” is
mostly nonsense, but I am trying to convey that a there is a tracer
flux even without any external tracer concentration difference.