This blog has a series on “sorption”,1 which I recommend everyone to read. A key message is that the mobility of exchangeable cations is not broadly internalized by the bentonite scientific community. I write
[I]t appears as the implication of releasing the immobilization assumption has not been fully comprehended. To this day, the traditional model is used as the basis for interpretation in basically all publications, to the extent that “\(D_a\)”, “\(D_e\)” and “\(K_d\)” usually are considered to be experimental quantities.
Recently I’ve come across a publication that illustrates these problems well: “Influence of temperature and dry density coupled effects on HTO, 36Cl, 85Sr and 133Ba diffusion through compacted bentonite” (García-Gutiérrez et al., 2024). Here I will review this paper, as part of my aim to review already published papers. We refer to the study in the following as GG24. It should be emphasized that this paper is merely an illustrative example of problems that I mean are found everywhere in the compacted bentonite research field.
Overview
As the title reveals, the main investigation of GG24 concerns diffusion of various aqueous species (HTO, Sr, Br, and Cl). Measurements were done in samples of “FEBEX” bentonite at densities 1.2, 1.4 and 1.65 g/cm3, and at temperatures 25, 40, 60 and 80 °C. The tests used the so-called “planar source method”, which is performed in closed cells (i.e. no contact with external solutions). Apart from the diffusion results, GG24 also report “batch sorption experiments” for strontium and barium, where “FEBEX” bentonite was suspended in solutions containing the radionuclides. In addition, GG24 report tests where samples of density 1.65 g/cm3 were supposedly equilibrated with an external reservoir of “ultrapure water”, at different temperatures.
Apparent diffusivity has not been determined
GG24 claim without hesitation that they “determine” apparent diffusion coefficients of the various species. I mean that such diffusion coefficients are not what has been measured/determined. But, rather than me spending blog space to flesh out my arguments from scratch, and which I have already done a long time ago, let’s discuss this issue using the actual values reported in GG24 as a starting point.
Anyone will agree on that what has actually been measured in GG24 are concentration profiles in samples of compacted bentonite. The samples were prepared with the different radionuclides initially present as a thin central layer (the “planar source”). After allowing these radionuclides to diffuse for a specified duration, the samples were sectioned and analyzed. This procedure provided a quite large set (48) of concentration profiles for the different species at different densities and temperatures.
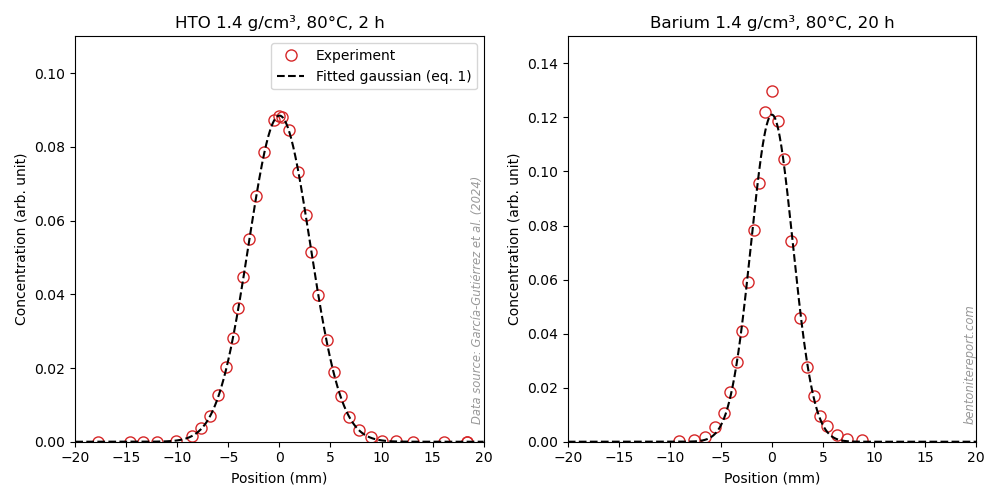
As the above figure shows, the concentration profiles demonstrably fit a Gaussian distribution
\begin{equation} c(x,t) = \frac{n_0}{\sqrt{4Dt\pi}}\exp\left (-\frac{x^2}{4Dt} \right ) \tag{1} \end{equation}
where \(t\) denotes time, \(x\) is the position relative to the initial central source, and \(n_0\) and \(D\) are constants. We will refer to \(D\) as the diffusion parameter, to avoid associating it with any particular model. Of course, eq. 1 is a solution to the 1-dimensional diffusion equation with an instantaneous point source (i.e., at \(t=0\), the total amount diffusive substance \(n_0\) is all assumed to be located at \(x=0\)). Without alluding to any particular model, we can thus safely conclude from the results presented in GG24 (and in many other studies) that concentrations in these types of systems evolve in accordance with a diffusion equation.
Apparent diffusivity of cations?
However, GG24, and most other researchers in this field, automatically interpret the diffusion parameter as an apparent diffusivity, i.e. that \(D= D_a\). Here is where I think a lot of problems start. To begin with, I get the feeling that many researchers are not fully aware that they make this extra step; I don’t know of how many papers I’ve read, where authors claim to measure or determine apparent diffusivities, as if these are direct experimental quantities.
To proceed, we must make clear what authors typically mean by the term “apparent diffusivity”. Here, researchers may be quite precise in one sense, while simultaneously — and seemingly unknowingly — blurring the definitional lines. GG24 provides a good example of this. The only model relation explicitly stated is the commonly encountered relation between apparent diffusivity (\(D_a\)), effective diffusivity (\(D_e\)), and distribution coefficient (\(K_d\)) (eq. 2 in GG24)
\begin{equation} D_a = \frac{D_e} {\phi + \rho_d K_d} = \frac{\phi D_p} {\phi + \rho_d K_d} \tag{2} \end{equation}
where \(\rho_d\) is dry density and \(\phi\) the physical porosity. In the second equality above we have also written the effective diffusivity in terms of the pore diffusivity (\(D_p\)), \(D_e = \phi D_p\).2
Indisputably, eq. 2 is a statement of the traditional diffusion-sorption model. Adopting this relation, GG24 consequently assume that:
- The entire pore volume is filled with bulk water, where ions diffuse
- Cations in this pore water is taken up, via ion exchange, by the solid and become immobile
All quantities in eq. 2 also have distinct physical interpretations. In particular:
- \(D_p\) (or \(D_e\)) is the real diffusion coefficient, i.e. \(D_p\) reflects the molecular mobility of the considered species within the porous system
- \(D_a\) reflects the combined effect of cations diffusing (\(D_p\)) and sorbing/becoming immobile (\(K_d\)) — hence the name “apparent”.
When GG24 claim that they determine \(D_a\) they are consequently saying that they observe a process where ions diffuse in a bulk water phase, while being retarded due to sorption. This is also explicitly stated (my emphasis)
A comparison of the diffusion behavior of HTO and 133Ba at the same dry density and temperature reveals that the distance traveled by 133Ba is consistently lower than that of HTO. This suggests that the diffusion time needed for the sorbing element to develop concentration profiles as wide as those observed for the non-sorbing element is much higher (ranging from 2 to 72 h), in agreement with the expected retardation due to adsorption in the clay.
I’m quite astonished to see researchers make such a statement in 2024. Sure, I nag constantly on the blog about flawed conceptual models of compacted bentonite that refuse to die. But I have been under the impression that it at least was established that simple cations are not retarded in this way: various observations of “\(D_e\)” (or “\(D_p\)”) being greater than the corresponding diffusivity in pure water have been reported for a long time; most model development for the last 40 years or so has been attempts to deal with exchangeable ions being mobile; and terms like “surface mobility” seem rather established.
But even if authors in the contemporary bentonite scientific literature occasionally acknowledge some type of mobility of “sorbed” cations, it is simultaneously clear that the conceptual ideas underlying the traditional diffusion-sorption model has never been eradicated. I find this quite frustrating and sad, especially since several research groups drew correct conclusions already in the early 1980s, as I discuss here. This is e.g. what Jahnke and Radke wrote in 1985(!)
[…] sorption does not reduce the effective diffusion coefficient within compacted packing much at all. This implies that \(K_d\) values are irrelevant during the transient release period. That is, we do not anticipate large reductions in transient ion migration rates due to sorption retardation on montmorillonite.
[…]
\(K_d\) values are of extreme importance during the steady release period. Counter to what may have been anticipated, larger \(K_d\) values lead to higher release rates.
Apart from all empirical evidence of mobility of “sorbed” cations, and the fact that the traditional diffusion-sorption model (eq. 2) makes completely erroneous assumptions about compacted bentonite, we can use the actual values reported in GG24 to debunk their statement that cations are retarded due to adsorption. Solving eq. 2 for \(D_p\) gives
\begin{equation} D_p = \left ( 1 + \rho_d K_d/\phi \right ) \cdot D_a \tag{3} \end{equation}
From the performed “batch sorption tests”, GG24 report separately evaluated values of \(K_d\), for the same aqueous chemistry as for the diffusion tests (the aqueous chemistry will be discussed further in the second part of this review). For barium at room temperature, GG24 determine \(K_d \approx\) 80 cm3/g. The table below shows calculated values of \(D_p\), using eq. 3 and this \(K_d\) value, for the reported Ba diffusion tests performed at 25 °C.
| \(\rho_d\) (g/cm3) | \(\phi\) (-) | \(D_a\) (m2/s) | \(D_p\) (% of \(D_0\)) (m2/s) |
| 1.20 | 0.556 | 1.50⋅10-11 | 2.6⋅10-9 (307%) |
| 1.40 | 0.481 | 0.95⋅10-11 | 2.2⋅10-9 (261%) |
| 1.65 | 0.389 | 0.36⋅10-11 | 1.2⋅10-9 (144%) |
For all tests, the “pore diffusivity” is much larger than the corresponding diffusion coefficient for barium in pure water (\(D_0\) = 8.5⋅10-10 m2/s), which of course is completely untenable. This inevitably demonstrates that “\(D_p\)” is not a pore diffusivity, and that “\(D_a\)”, consequently, is not an “apparent” diffusivity.
Doing the same exercise for the strontium data in GG24 (\(K_d \approx\) 10 cm3/g) does not give “pore diffusivities” larger than those in pure water, but in the range 65% — 27% of \(D_0\). This does of course not mean that the traditional diffusion-sorption model applies to strontium!3 The evaluated values for “\(D_p\)” are still unrealistically large, and we know from many other experiments that “sorbed” Sr is not immobile. In fact, strontium diffusion in dry bentonite was an early observation demonstrating that the traditional model is invalid. We must also recognize that it only takes the counterexample of a single type of ion to invalidate the model as a whole — bentonite cannot contain only bulk water for only some ions.
Apparent diffusivity of chloride?!
GG24 furthermore claim to also “determine” “apparent” diffusivity of chloride. But this is not reasonable even from a model point of view. Chloride is described as “non-sorbing” (\(K_d = 0\)), so even if this ion could be modeled using the traditional diffusion-sorption model, it is quite misleading to claim that the “determined” diffusivity is apparent; for a “non-sorbing” species, eq. 2 directly shows that \(D_a = D_p\), i.e. that the “apparent” diffusivity reduces to the pore diffusivity. To report “apparent” diffusivity in this case is a little like insisting on stating the gross weight of a consumer good that has no packaging.
But the problems are deeper than simply failing to properly identify the nature of the diffusion parameter, because the traditional diffusion-sorption model is not able to describe anion diffusion. Specifically, an overwhelming amount of empirical evidence shows that \(D_e \neq \phi D_p\) for anions. To at all speak of “apparent” diffusivity of anions is thus to apply an inapplicable model concept. Many researchers don’t seem very bothered by this, and quite happily continue to apply concepts and parameters of the traditional diffusion-sorption model also to anions. But, in doing so, they typically, and seemingly unknowingly, switch underlying conceptual model on the fly. For example, GG24 write
Despite being a non-sorbing species, the transport of 36Cl differs from that of HTO in compacted clays due to the impact of electrostatic forces between the negatively charged clay surfaces and the anions. This leads to anionic exclusion, which reduces the volume of pore water available for anion transport and results in smaller values of \(D_a\) compared to those of HTO.
Leaving aside for the moment that this is not how anion exclusion works, note how the conceptual view is suddenly changed: the pore volume is now assumed to have different parts, with distinctly different properties. Researchers often brushes off this complication by simply replacing the porosity parameter in the traditional diffusion-sorption model with an “effective porosity”, \(\epsilon_\mathrm{eff}\). But this is of course not allowed! The physical porosity appears in eqs. 2 and 3 because the traditional diffusion-sorption model distinctly assumes all pore water to be bulk water. The pore water cannot all be bulk water at the same time as it is not. In describing anions, researchers “silently” actually switches to the effective porosity model.
The effective porosity model assumes that some of the pore water is bulk water, quantified by \(\epsilon_\mathrm{eff}\), while the rest has non-specified non-bulk properties. This is
incompatible with the traditional diffusion-sorption model.4 The effective porosity model, moreover, doesn’t include sorption, and consequently has no concept of an apparent diffusivity (i.e. it does not describe a combined process of ions diffusing and being taken up by the solid). The governing equation for the effective porosity model is a diffusion equation with a pore diffusivity as parameter, i.e. \(D = D_p\) in eq. 1.
We have thus seen that the diffusion parameter (eq. 1) cannot be interpreted as an apparent diffusivity for cations. For anions, the apparent diffusivity concept does not even apply, and the actual invoked model relates the measured concentration profile with a pore diffusivity. It follows that under no circumstances is it adequate to state that apparent diffusion coefficients are determined in these types of tests.5 This goes for GG24, as well as for many many other studies within the bentonite research field.
We now must ask if the effective porosity model is appropriate for describing bentonite. Note how far out in the weeds we are: we — not the researchers doing the reporting — have first identified that it actually is the effective porosity model that is used (for chloride), and subsequently have to ask if this model is valid. The short answer is that this model does not describe compacted bentonite.
To elaborate on this, we note that GG24 undoubtedly assume that the effective porosity is physical, as they write “This leads to anionic exclusion, which reduces the volume of pore water available for anion transport and results in smaller values of \(D_a\) compared to those of HTO.”6 But there is vast evidence that this is not a valid interpretation.
If anions were restricted to a smaller part of the pore volume, it is reasonable to assume that their pore diffusivity also decreases with the parameter \(\epsilon_\mathrm{eff}\), as GG24 imply. This is, for example, the rationale behind Archie’s law,7 which assumes that pore diffusivity depends on porosity as \(D_p = \phi^n D_0\) with \(n>0\), and \(D_0\) being the diffusivity in pure bulk water. Also, for a porosity approaching zero, the diffusivity may be expected to show a threshold behavior, as the pore volume may become disconnected at some specific porosity or in some specific porosity interval. Under any reasonable circumstance, if effective porosity quantifies an actual physical pore volume, we can be certain that \(D_p\) should decrease with decreasing effective porosity, and approach zero in the limit of vanishing effective porosity. But this is not what is observed.
Variations with ionic strength are usually explored in through-diffusion experiments, and data from such tests are often quite uncertain, as we noted in the assessment of chloride equilibrium concentrations. But the results from e.g. Van Loon et al. (2007) show unambiguously that the chloride pore diffusivity does not change significantly, as the ionic strength is varied by two orders of magnitude (with corresponding significant changes of the effective porosity parameter).8
However, the most clear-cut evidence invalidating the effective porosity model is obtained from closed-cell diffusion tests performed at (near) zero ionic strength. In pure montmorillonite samples saturated with distilled water, the effective porosity parameter is minute, perhaps even strictly zero. Yet, e.g. Kozaki et al. (1998, 2001) consistently measure chloride pore diffusivities in such systems that show no signs whatsoever to be influenced by such a minimal effective porosity.
Summary
GG24, and most other researchers in this field, claim to “determine” apparent diffusivities, for both cations and anions, in these type of tests. For cations we can use the data provided in the article to clearly demonstrate that what has been measured are not apparent diffusivities. For (simple) anions, the concept of apparent diffusivity does not even apply and GG24 actually interpret these results in terms of the effective porosity model (perhaps without being fully aware of it). The effective porosity model, moreover, can clearly be demonstrated to not be valid for bentonite.
We are thus in the following quite awkward situation:
- An entire research community is wrong about what transport processes are actually active in compacted bentonite.
- At the same time, many ion distributions undoubtedly evolve in bentonite in accordance with the diffusion equation.
However, to simply state that the research community is wrong does not fully describe the situation. Even if researchers routinely use terms such as apparent diffusivity and effective (or anion-accessible) porosity, and make definite statements about mechanisms (as exemplified above), it appears to me that they simultaneously often do not fully “believe” their own statements. Rather, it seems to me to be kind of “mandatory” in bentonite diffusion publications to regurgitate statements relating to the traditional diffusion-sorption and effective porosity models, while “\(D_a\)”, in practice, is simply the name given for the diffusion parameter.
In the next part of this review we will continue to examine the conceptual confusion in GG24, in particular with respect to the term “pore water”, and how the observed temperature dependencies are discussed.
Footnotes
[1] “Sorption” is written with quotation marks here, because the message is that “sorbing” ions typically do not become immobilized, in contradiction with the traditional diffusion-sorption model.
[2] Just as the word “apparent” is supposed to have meaning, “effective” originally refers to scaling the flux to the entire cross sectional area of a porous system.
[3] We may also consider the values of \(K_d\) reported in GG24 for “deionized water”. In this case, \(K_d\) at room temperature is above 5000 cm3/g for Sr and above 8000 cm3/g for Ba. If we entertain the idea that the “pore diffusivities” evaluated with “pore waters” actually are pore diffusivities, this implies that GG24 predict that values of “\(D_a\)” in systems equilibrated with pure water should be of the order of 10-13 — 10-14 m2/s, i.e. that they would have to perform the tests for several months (!) to get concentration profiles comparable to those reported. I’ll stick my head out and claim that such values of “\(D_a\)” will not be measured for barium in “FEBEX” samples equilibrated with pure water. Rather, “\(D_a\)” in such systems will be comparable to what GG24 measure in “pore water” (Because “\(D_a\)” is essentially independent of ionic strength). This, in turn, implies that “\(D_p\)” for barium in samples equilibrated with pure water will be of the order 10-7 m2/s, or about 100 times larger than bulk water diffusivities!
[4] To reconcile the two models, you may for instance modify the traditional diffusion-sorption model by assuming that the bulk water phase only occupies a part of the pore space. But this would mean e.g. that calculated values of \(D_p\) would become even more preposterous (\(\epsilon_\mathrm{eff}\) replacing \(\phi\) in eq. 3). It also makes the description much less coherent, as the non-bulk part is left completely passive and non-described, as “sorption” still is associated with the solid. You may then of course try to reconcile that, by instead ascribing the “sorption” to the non-bulk part of the pore space (perhaps labeling this part the “diffuse layer”?)… You have now stepped into the multi-porous model swamp. This regression simply demonstrates that the traditional diffusion-sorption model is not a reasonable starting point for modeling compacted bentonite.
[5] This statement refers strictly to simple ions. There are also aqueous species that truly sorb, on montmorillonite edges or other types of minerals that may be present in the bentonite.
[6] As we have now learned, this statement (which still is wrong!) should be about \(D_p\), not \(D_a\).
[7] I have just realized that I use the term Archie’s law quite sloppily on the blog. The authors that originally (as far as i understand) came up with this description of diffusivity in porous systems suggested an analogy with the actual Archie’s law, which describes electrical resistivity. However, in order to keep the amount of words down a little, I will continue to use the term Archie’s law also for diffusivity. This is also how it is typically used nowadays in the bentonite scientific literature.
[8] Earlier, we have also demonstrated that the effective diffusivity in Van Loon et al. (2007) does not obey Archie’s law